Biomass as Congealed Electricity

Part Of: Biology sequence
Followup To: A Portrait of LUCA
Content Summary: 5800 words, 29 min read

Redox Chemistry

Last time, we reconstructed the nature of LUCA, the root at the tree of life. Today, we’re going to be taking a deeper look at how life makes a living: how cells harvests electricity to build more of themselves.

Life requires energy influx. Energy influx creates disequilibria and maintains them against relaxation. Some disequilibria are differences between locations. A temperature gradient, a pressure difference, and a concentration difference across a membrane are spatial disequilibria, and they relax by transport. Other disequilibria are differences between the present state of a system and an equilibrium state it can reach without moving anywhere. A supersaturated solution, a mixture of hydrogen and oxygen, and an activated molecule such as ATP or a thioester each hold a state disequilibrium, and they relax by transformation.
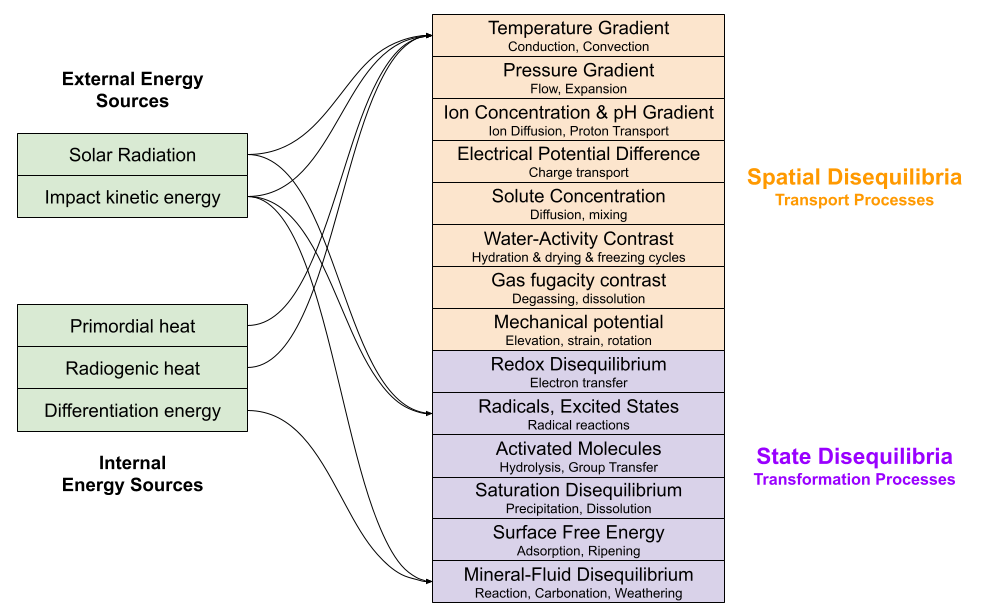
Out of this menu of fourteen options, life runs a state disequilibrium called redox.  Oxidation is electron loss, reduction is electron gain (recall the mnemonic OIL RIG). Redox (reduction-oxidation) reactions involve both of these as electron transfer dissipates this disequilibrium. 

The remainder of this post will explore how life uses redox, and where these disequilibria come from.

A redox couple is the oxidized and reduced form of the same chemical species. When hydrogen gas (H₂) is oxidized, its bond breaks and it releases two electrons, becoming 2H⁺. When carbon dioxide (CO₂) accepts eight electrons (along with eight protons), it is ultimately reduced to methane (CH₄) and water. A donor-acceptor pairing combines two such half-reactions. We write this pairing as 2H⁺/H₂ → CO₂/CH₄. 

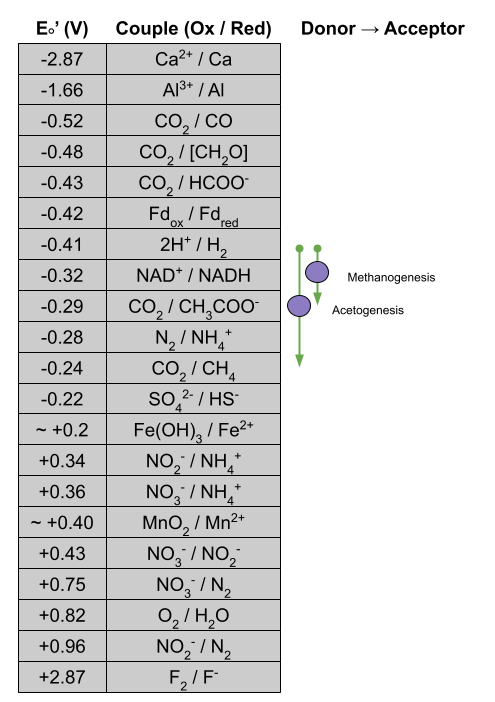
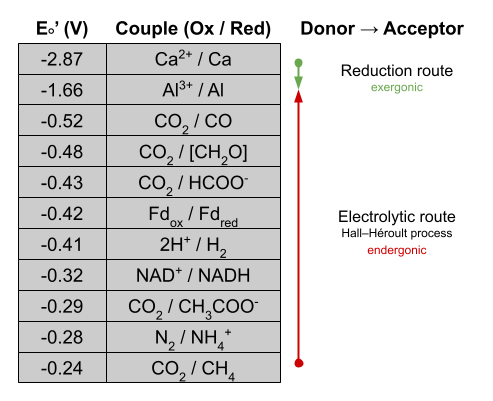
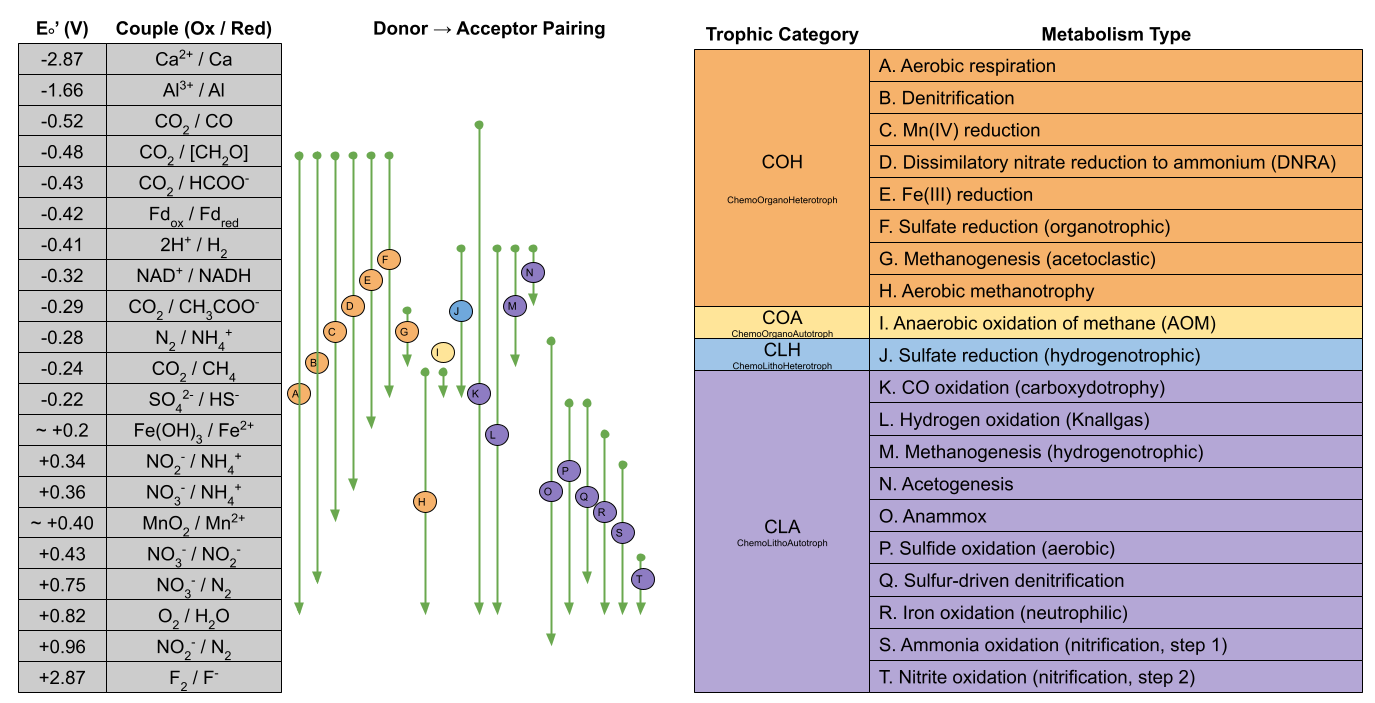
Each couple has a characteristic voltage known as the reduction potential (Eo’). Strongly negative values indicate a couple that gives electrons up readily; positive values represent willingness to accept. Spontaneous donor-acceptor pairings run from smaller reduction potentials to larger. The potential difference ΔE°′ = E°′(acceptor) − E°′(donor) must be positive. Since 2H⁺/H₂ sits at −0.41 V, and CO₂/CH₄ is −0.24 V (potentials at pH 7 from Thauer et al 1977), a methanogen running from hydrogen gas to carbon dioxide gets ΔE°′ = (-0.24) – (-0.41) = 0.17 V. Actual potentials shift with concentrations, gas pressures, and pH, so the available energy depends on the organism’s conditions. 

These relationships are often represented with a redox tower. This is done by sorting couples by reduction potential, and inverting the y-axis. The low reduction potentials (strong donors) sit at the top, and the high values (strong acceptors) are at the bottom. Spontaneous donor-acceptor pairings with positive potential difference can be represented as arrows cascading down the tower. Importantly, the magnitude of the potential difference (arrow length) determines the energy released: ΔG°′ = −nFΔE°′, where n is the number of electrons transferred and F is Faraday’s constant.
Electrochemical Cells

Redox disequilibria relax by transformation: electrons transfer in situ. An electrochemical cell separates the two half-reactions in space, connecting a state disequilibrium (the redox gap) to a spatial one (a voltage across space). Oxidation (electron donation) takes place at the anode, reduction (acceptance) takes place at the cathode. Electrons travel from anode to cathode through a conductor. The accumulating charge differences would quickly squelch the redox reaction. To achieve a cell rather than a capacitor, an electrolyte (e.g., salt solution) carries ionic current between the reaction sites. This closes the circuit and balances charge. 

We might distinguish between two forms of cell. In a galvanic cell, the donor-acceptor pairing is spontaneous (exergonic). Galvanic cells convert chemical potential into work. They “discharge the battery.” In an electrolytic cell, the donor-acceptor pairings are non-spontaneous (endergonic). These cells consume work to generate chemical potential. They “charge the battery.” 

We will soon learn that biological cells are electrochemical cells. To better contextualize this fact, we first turn to non-biological examples. 

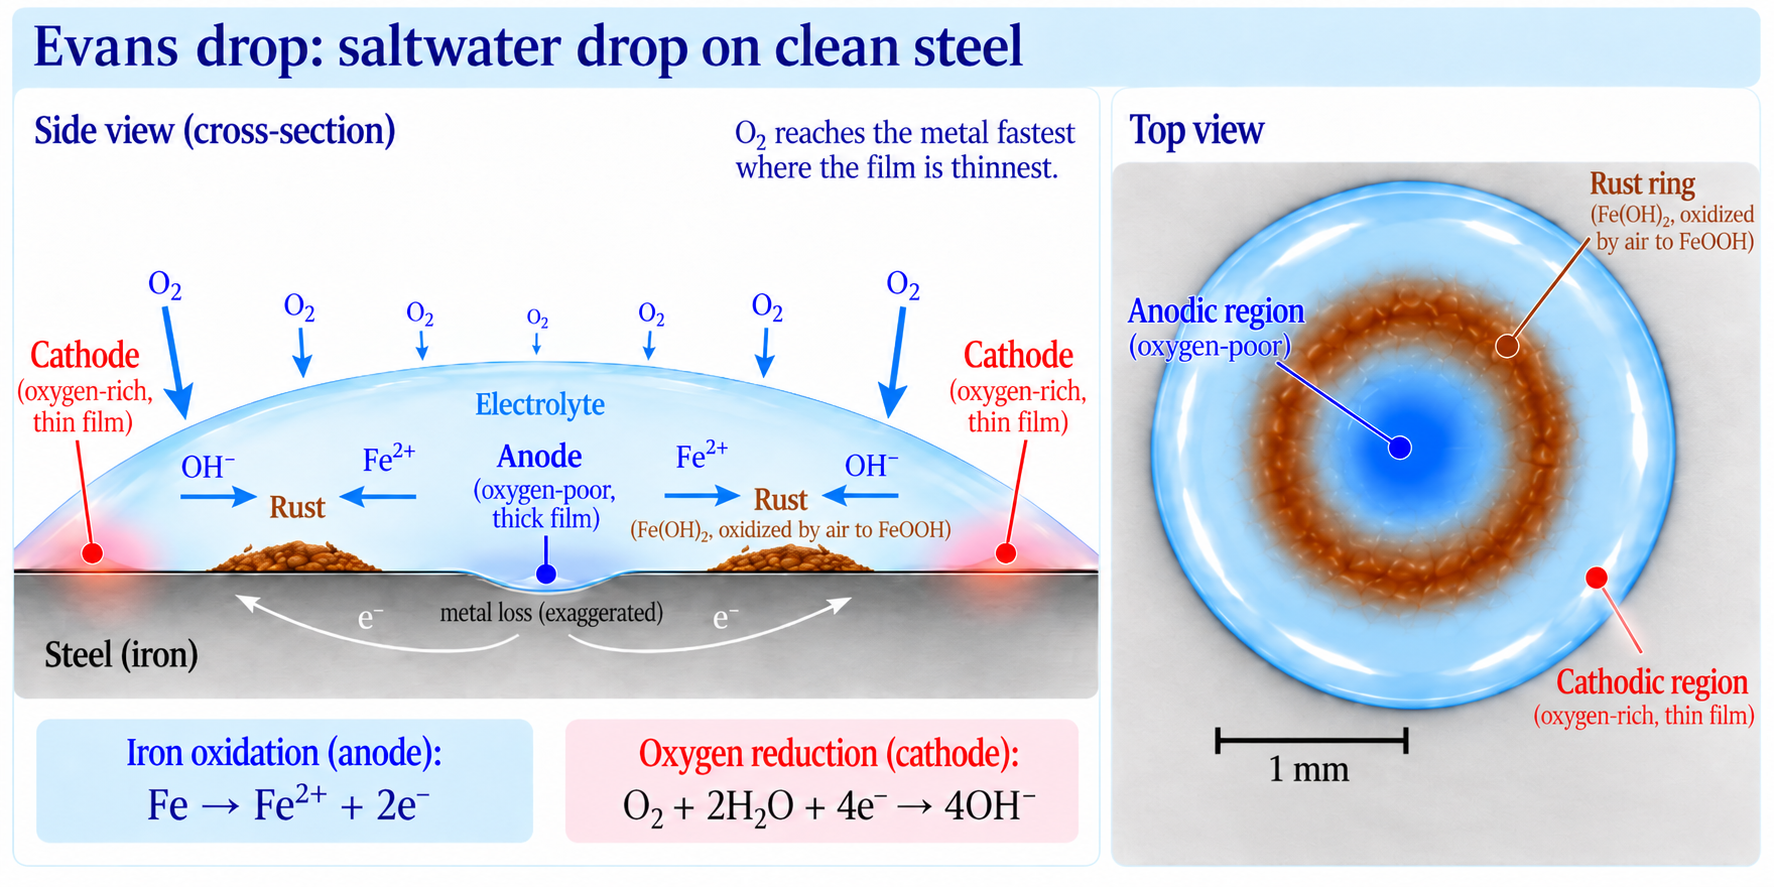
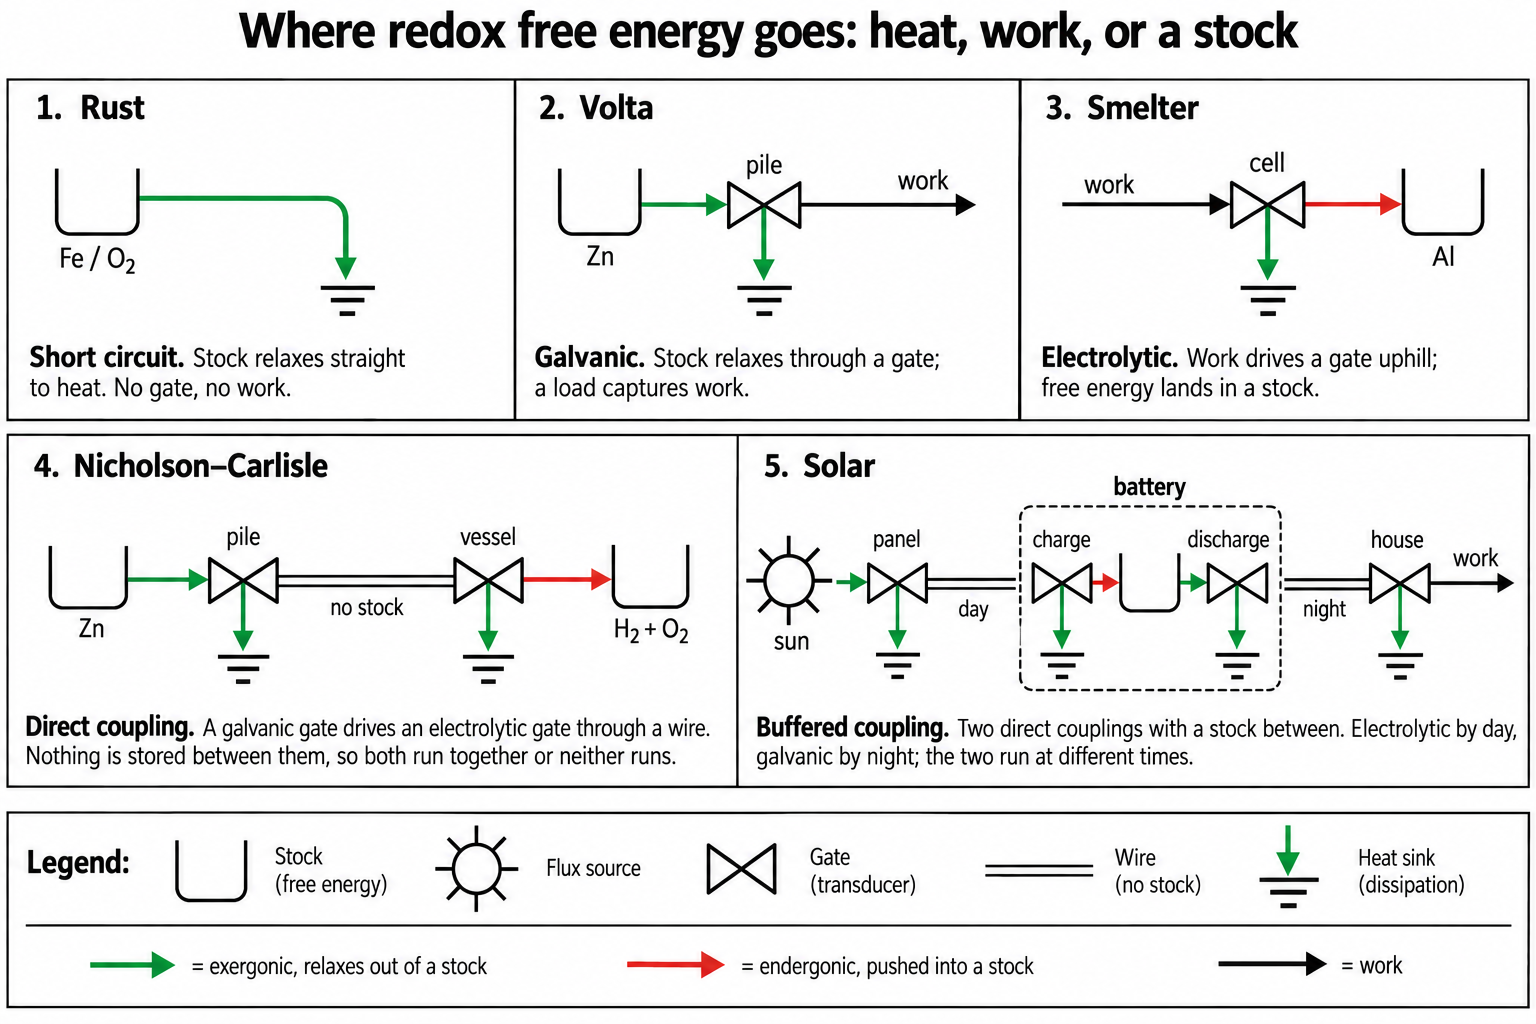
Consider the Evans experiment, a single drop of water placed on clean steel. Oxygen reaches the metal fastest near the border of the drop, where the film is thinnest, and slowest at the center. The oxygen-poor center becomes the anode, where iron is oxidized to Fe²⁺ and releases electrons. The oxygen-rich periphery becomes the cathode, where dissolved oxygen accepts those electrons and is reduced to OH⁻. Fe²⁺ and OH⁻ diffuse towards each other and precipitate, producing a characteristic ring of rust. These corrosion processes are ubiquitous because they are exergonic and, on iron, unburdened with a protective oxide film. But here, electron flow is effectively short-circuited through the metal itself. The available Gibbs free energy is dissipated mainly as heat. 
This same process can be harnessed to do work. Alessandro Volta opened the short circuit in 1800. His voltaic pile consisted of repeated zinc–electrolyte–copper sandwiches, each forming a galvanic cell.  Zinc oxidation releases electrons, which are consumed by a reduction at the copper electrode. Because the electrodes are separate pieces of metal, the electrons cannot pass directly between them. They must travel through an external wire, and anything placed in that path, which we will call the load, can capture part of the redox free-energy drop as useful work.

The pile runs because zinc oxidation is downhill. But many reactions we want run uphill. Aluminum is a prized metal, yet on Earth nearly all of it is bound to oxygen. Smelters convert aluminum oxide (Al₂O₃) into aluminum metal by adding three electrons: Al³⁺ + 3e⁻ → Al. Aluminum is a very weak oxidant, with a −1.66 volt potential. Alumina can be reduced with a stronger reductant. Deville did this in 1854 with sodium metal. But the sodium was difficult to manufacture, and aluminum production was as expensive as silver. The Hall–Héroult process lowered costs by instead supplying reducing power directly from electricity. Alumina is dissolved in molten cryolite, and a large current drives the reduction of Al(III) at the cathode to liquid aluminum while carbon is oxidized at the anode. This reaction is non-spontaneous without an external power source. This is why the industry calls aluminum congealed electricity. The metal is a stock: a reservoir of free energy, filled by one process and drained later by another. 
What provides work to an electrolytic cell? A galvanic cell can. Within weeks of Volta’s discovery, Nicholson and Carlisle placed the pile’s wires in water and watched hydrogen and oxygen appear. The voltaic pile supplied the 1.23 volts needed to decompose water. The wire connects driving and driven processes. This wire cannot sustain the reaction when the battery depletes. Both processes must proceed in tandem. In direct coupling, a driving process and a driven process are joined without enough storage to separate their operation in time. 

Direct coupling is insufficient when energy inflow and outflow proceed at different times and different rates. Consider an off-grid solar installation. During the day the panels supply electrical work, but a house draws power continuously. This setup is enabled if a stock is placed on the wire. By day, the stock (a battery) is charged in an electrolytic process. But whenever the house needs power, the battery discharges in a galvanic process. In buffered coupling, a stock separates two processes in time. The stockis filled when power is available and depleted when it is needed. A single cell can perform both electrolytic and galvanic roles. 
Respiration as a Galvanic Cell

Humans eat food and breathe oxygen. Why? Both are delivered to our cells where electrons are stripped from organic matter and sent down the respiratory chain to oxygen. Cellular respiration is a donor-acceptor pairing. Life runs on redox chemistry. 

Consider Cupriavidus necator, a bacterium that strips electrons from hydrogen to give to oxygen. We’ve already seen the electrolytic cell from the Nicholson-Carlisle experiment, which consumed energy to separate water. In 1839, William Grove constructed the reverse. He placed platinum electrodes in separate tubes of hydrogen and oxygen, connected them through an acid, and observed a current. Cupriavidus harvests energy in precisely the same way as Grove’s device: 2H⁺/H₂ against O₂/H₂O. But life utilizes this energy differently, as we will see below. 

Respiration occurs in the electron transport chain (ETC). While the modern ETC has many components, for our purposes we consider three. Electrons are passed from the entry enzyme towards the exit enzyme, which function as anode and cathode respectively. 

The entry enzyme and the exit enzyme differ across organisms. The quinone pool represents their interface. The entry enzyme (e.g., a membrane-bound hydrogenase) must accept electrons from a redox donor and reduce quinone. Quinone takes the two electrons and, from the inner face, two protons, becoming quinol. The exit enzyme (e.g., quinol oxidase), needs only oxidize quinol. (In mitochondria an extra stage, Complex III and cytochrome c, sits between the pool and the exit enzyme.)

The journey of electrons occurs without a wire. Within the proteins, an electron will tunnel between fixed metal centers, iron-sulfur clusters and hemes, spaced no more than about 14 Å apart (Page et al 1999). Between proteins, the electrons continue their journey hitching a ride on the quinone. 

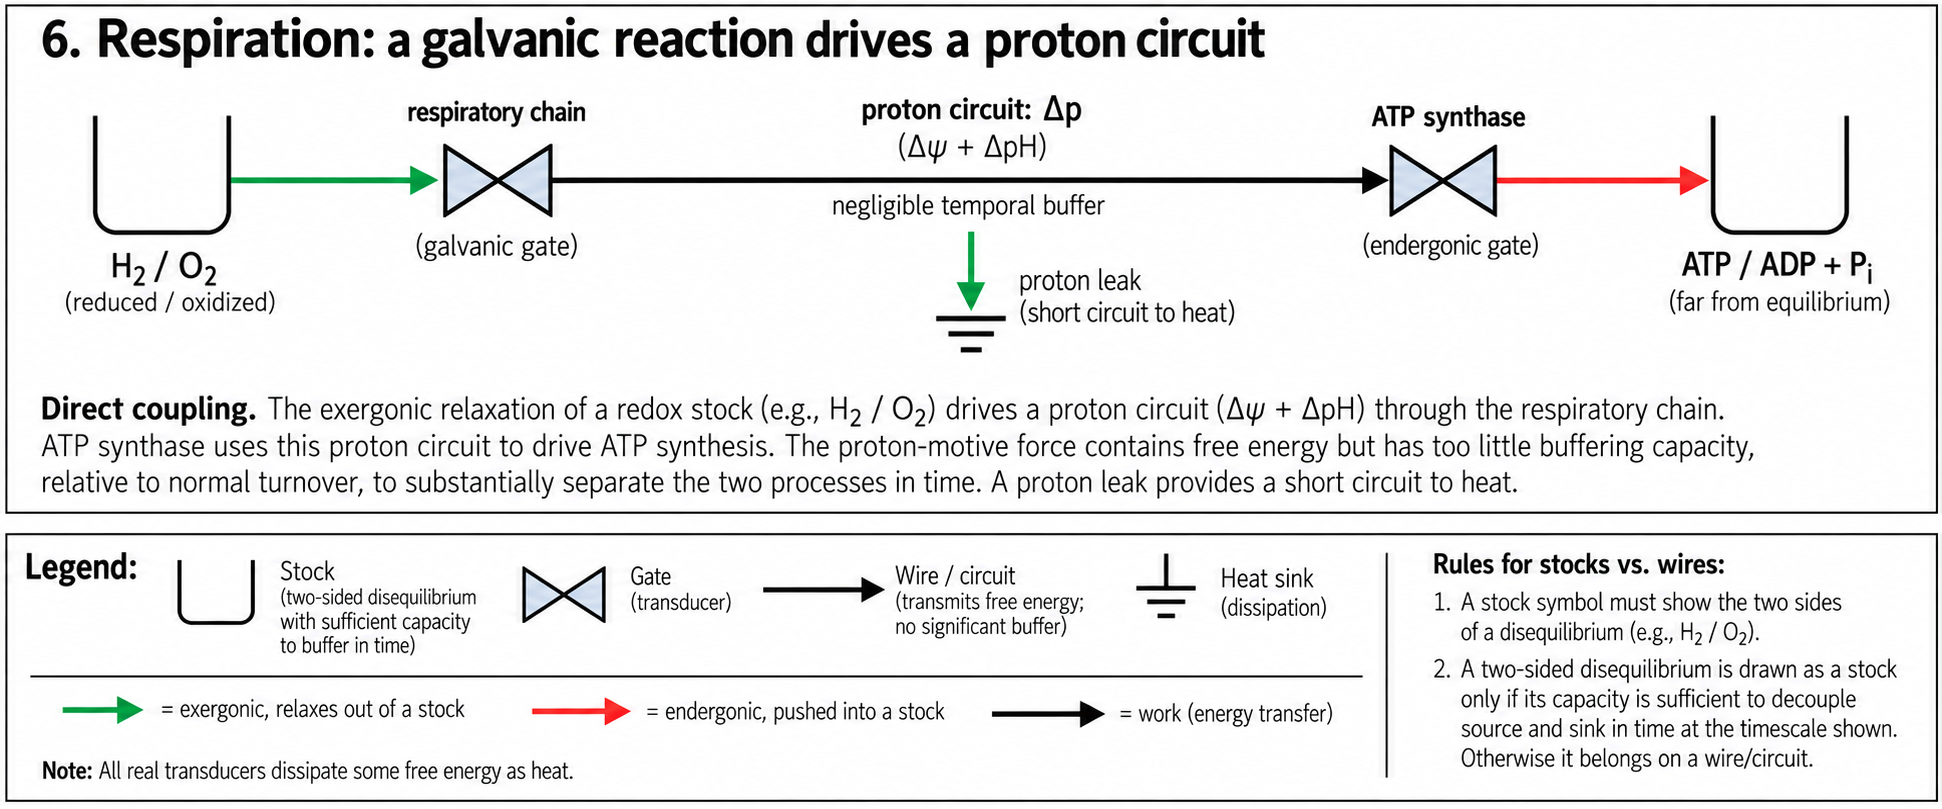
Where does the work go? The ETC enzymes are all embedded in a lipid membrane, which in prokaryotes separates the cytoplasm inside from the periplasm outside. The entry enzyme oxidizes the donor and releases its protons in the periplasm. Quinone is reduced in the membrane and takes its two protons from the cytoplasm. The exit enzyme oxidizes quinol and releases those protons on the periplasm, and reduces the acceptor with further protons drawn from inside. Every pair of electrons that journeys from anode to cathode carries several protons from inside to outside. Redox is a driving process; the protein gradient is a driven process. This coupling of state to spatial disequilibrium is called chemiosmosis

The gradient has an electrical part, Δψ, and a chemical part, ΔpH, and Mitchell (1961) combined them into a single quantity, the proton-motive force (PMF): Δp = Δψ − 59 mV × ΔpH at 25 °C. In mitochondria and most bacteria Δp is 150 to 220 mV, most of it electrical. That is roughly 15 to 20 kJ per mole of protons, and across a membrane 5 nm thick it is a field of about 4 × 10⁷ V/m, ten times the field at which air breaks down and lightning strikes (Lane 2005). Yet despite its strength, the PMF has limited storage capacity. If proton pumping stops while ATP synthesis continues, the gradient runs down. The PMF therefore serves mainly as the immediate link between driving and driven processes. This is direct coupling

ETC behavior resembles a voltaic pile. If you connect it to a load (the ATP synthase, which we will meet next), the PMF produces work. But what happens if you remove or disable the ATP synthase? In this open circuit, respiration slows because protons cannot return and Δp rises until the chain stalls against it. Chance and Williams (1955) starved the ATP synthase by withholding ADP. Respiration slowed to a crawl, only recovering when ADP was reintroduced. If the membrane develops a leak, this powerful voltage would be discharged in a short circuit. This is how brown fat warms newborns through a regulated leak channel, UCP1. An uncontrolled leak is more dangerous, thermodynamically equivalent to burning sugar via combustion. A short circuit releases the disequilibrium into waste heat, without powering the ATP-producing load.

The PMF produced by the ETC is only biologically useful if it performs work. That work happens at the ATP synthase. This rotor produces an activated molecule known as adenosine triphosphate (ATP). Thermodynamics favors hydrolysis of its terminal phosphate bond, leading to more stable adenosine diphosphate (ADP). Proton flux turns the synthase rotor, and the rotation forces ADP and phosphate together on the inner face. Three to five protons pass per ATP, depending on the organism. This process holds ATP, relative to ADP and phosphate, some ten orders of magnitude above the level equilibrium would allow. 

The ATP stock provides a buffer, but it is small. A human turns their whole ATP stock over within a minute or two, working muscle within seconds. The body regenerates roughly its own mass in ATP each day. ATP is called the energy currency of the cell. In many metabolic processes, ATP is used to enable non-spontaneous reactions by the group transfer mechanism. The terminal phosphate is handed to a substrate, which is lifted into a less stable state, and the endergonic step that follows is paid for by the relaxation of ATP toward ADP. Most endergonic steps in metabolism are paid this way. The rest are paid in reducing power, as we will see shortly.
Biological supply and demand rarely coincide. A plant captures sunlight by day but respires continuously. An animal eats at intervals and works continuously. A bacterium meets its substrate in pulses. Our solar-powered house solved this mismatch with the battery. Cells achieve the same buffered coupling solution with hierarchical storage. Phosphate reserves can become ATP directly. But a cell can dissolve only so much. Any reserve held in solution, like phosphocreatine in muscle, is shallow. Carbon-fuel reserves bypass this constraint by leaving the solution via polymerization (which cuts the particle count dramatically) or phase separation. Carbon-fuel reserves (including glycogen, starch, fat, and PHB) thereby unlock much deeper capacity and can last from hours to months. They rely on metabolic processes to convert their redox disequilibrium back into ATP.

We have sketched how redox disequilibrium is harvested to produce the PMF and ATP. But which molecular species does life harvest? 

Modularity and Habitat Colonization

The remarkable fact is that life does not care which span. Animals transfer electrons from organic carbon to oxygen. Prokaryotes can instead use donors and acceptors drawn from minerals, dissolved compounds, and gases. The same ETC (quinone pools, iron-sulfur clusters, a rotary ATP synthase) serves an astonishing diversity of habitats. The entry and exit enzymes are simply swapped. Life most likely began with a single donor-acceptor coupling. But modern life uses modular respiration. Donor and acceptor are configurable; the electron transport chain is largely conserved (Jelen et al 2016).

The voltage gap between an organism’s electron donor and acceptor sets the energy it can extract per electron. Power also requires current: a supply of donor and acceptor in the same place. Because life runs in water, the voltage gap has a ceiling. A substance strong enough to change the oxidation state of water will react with the solvent and cannot reach the organism. “Hydrogen gas (E°′ = −0.41 V) is the strongest reductant that accumulates in water, and O₂ (E°′ = +0.82 V) is the strongest oxidant that is both abundant and stable in it. As a stylized fact, the largest redox gap available in water is about 1.2 volts.

But on the Hadean Earth, oxygen was absent in bulk because nothing besides water photolysis produced it. The first bioenergetic revolution was the origin of life itself. The best electron acceptors available (carbon dioxide, -0.24 V; ferric iron, ≈ 0.0 V) sat within 0.5 volts of organic carbon. Limited by donor supply, the anaerobic biosphere could fix ~10¹³ mol C per year, or ~0.1 TW (Crockford et al 2023). 

Oxygenic photosynthesis heralded the second bioenergetic revolution. Cyanobacteria pushed electrons from water (+0.82 V) into carbon (-0.43 V). Water is inexhaustible, so primary production rose by an order of magnitude, to ~10¹⁴ mol C per year or ~1 TW. The waste product oxygen eventually accumulated in the atmosphere. Heterotrophs were then able to respire this span in the reverse direction. This second revolution allowed life to harvest near its maximum redox span. Subsequent revolutions had to come from current, by relaxing supply constraints.
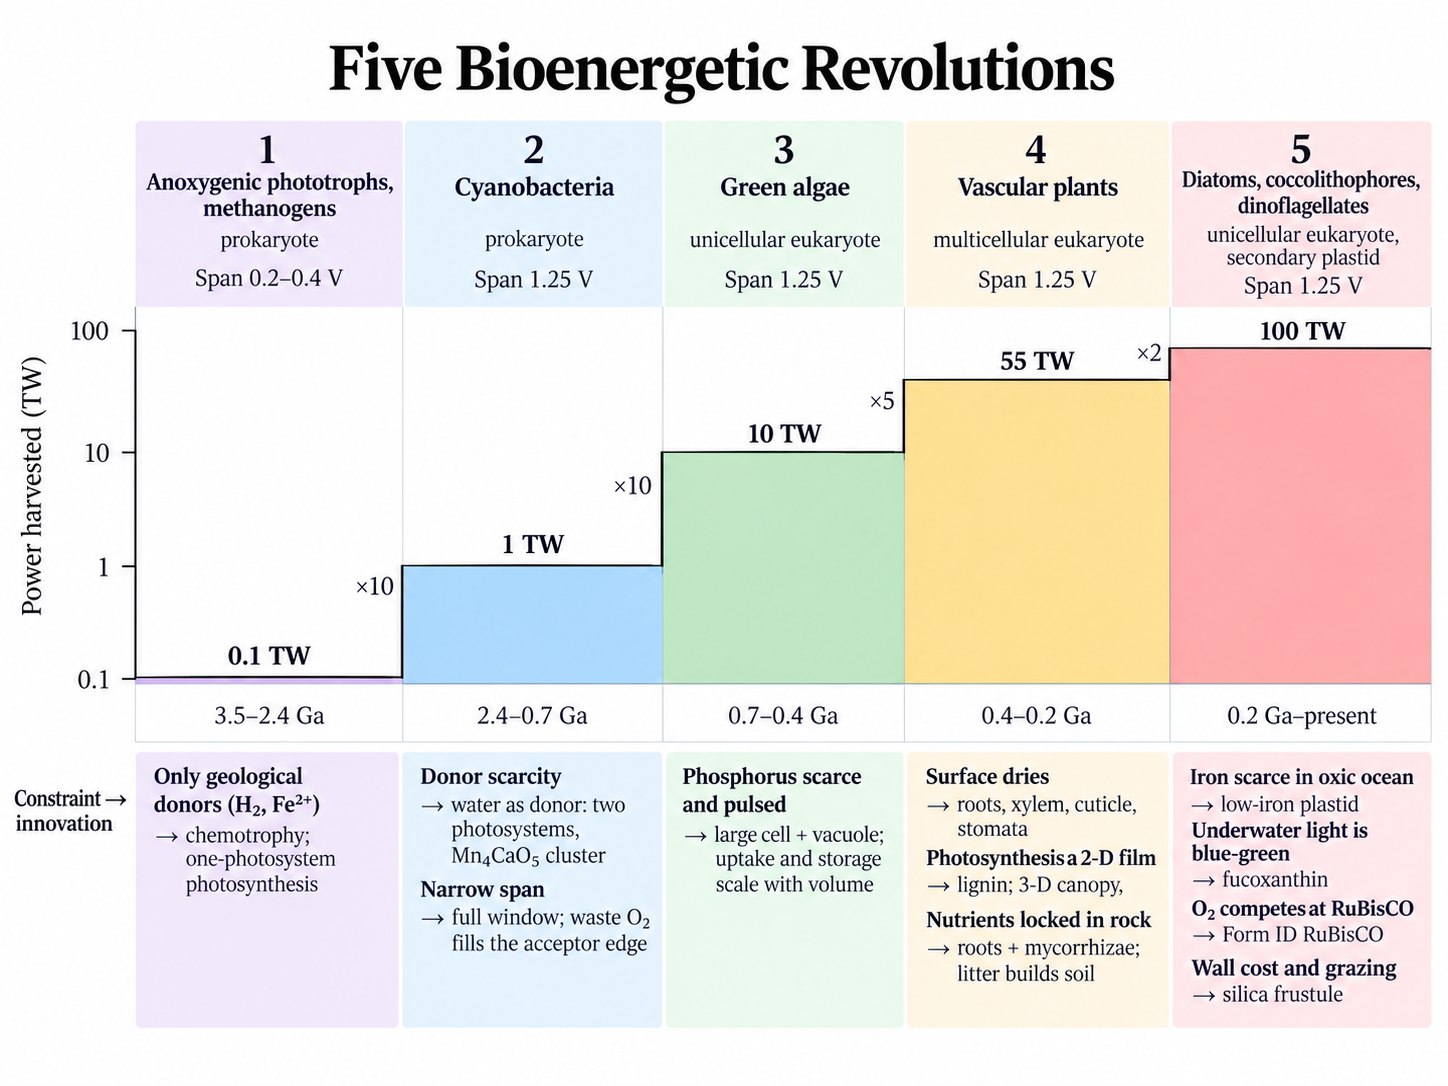
But life invaded myriad habitats in the early Earth even before oxygenic photosynthesis. The modularity of respiration allows for life to harvest nearly any form of redox disequilibrium anywhere.

A pair of electrons falling 1.23 V from H₂ to O₂ pays 237 kJ per mole of H2, while the same pair falling 0.17 V from H₂ to CO₂, the methanogen’s living, pays 33 kJ. Between them lie the denitrifiers, iron reducers, and sulfate reducers. Redox pairings correlate with sediment depth. Oxygen is consumed at the surface, followed by nitrate, manganese, iron, sulfate, then finally CO2 at the deepest layers (Froelich et al 1979). The biosphere fully occupies the redox tower (Falkowski et al 2008).

Microbiologists have even used gaps in the tower to predict the existence of new organisms. In 1977 Engelbert Broda listed a pairing the tower permitted but no known organism used: ammonium as donor against nitrite as acceptor. He argued that such lithotrophs should exist. They were found in the 1990s (Broda 1977; Strous et al. 1999). 
In sum, the biosphere runs galvanic cells to convert redox disequilibria into a stock of ATP. This is energy metabolism. What does a cell buy with it? 

The Machinery for Biosynthesis 

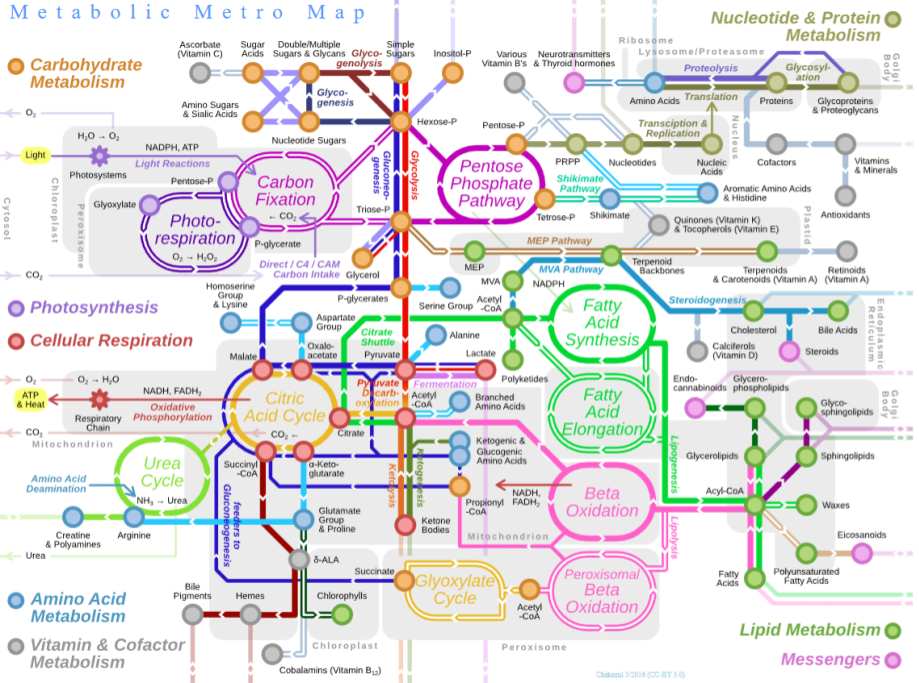
Life uses this energy to self-replicate, and making more cells requires synthesizing biomass. Energy metabolism finances carbon metabolism.

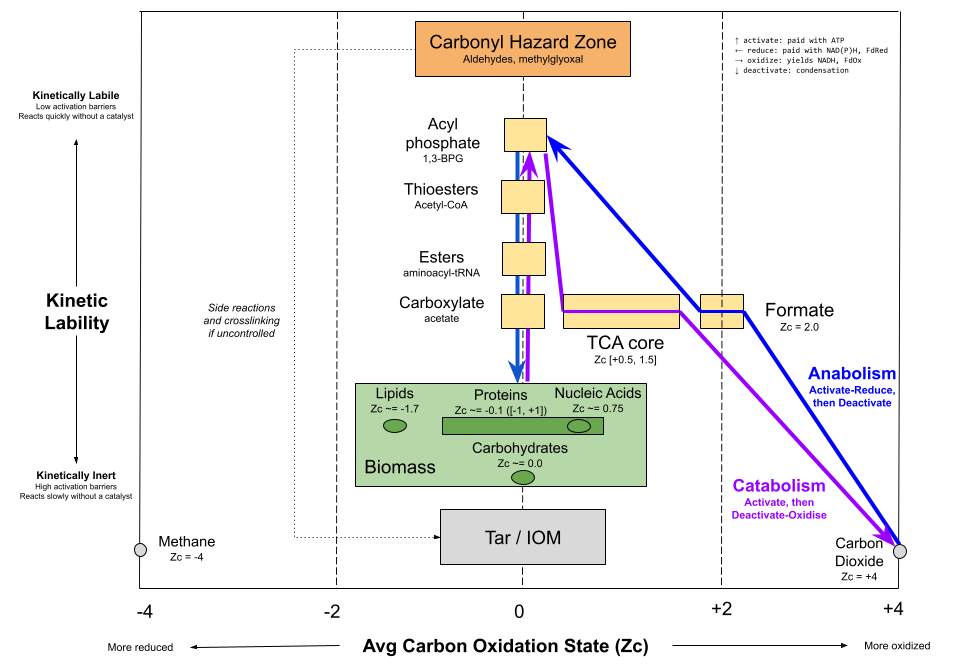
Let’s review how redox chemistry interacts with organic chemistry.  Autotrophs consume CO₂ and convert it into organic matter via carbon fixation. Carbon fixation transfers two electrons at a time from reductants to carbon. Each transfer moves a carbon atom one rung along a ladder. Define L as the total bond order from a carbon to oxygen, nitrogen, or sulfur, with a double bond counting twice.  As L decreases, the atom transforms from CO2 to carboxyl to carbonyl to alcohol to hydrocarbon. 
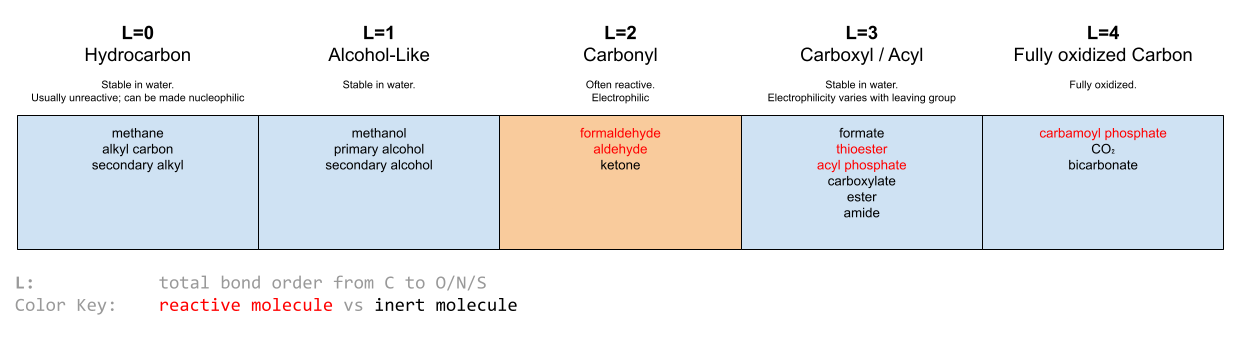
Losing L means reducing carbon. But redox does not uniquely constrain carbon chemistry. For bond formation, what matters is the local distribution of those electrons. An electrophile is an electron-poor atom that accepts electrons to form a bond, while a nucleophile is an electron-rich atom or group that donates it. Carbonyls expose a C=O bond, which is electrophilic. Hydrocarbons and alcohols do not expose this, while carboxylates at L = 3 stabilize theirs through resonance and negative charge. 

Biomass is deposited as alcohols and carboxyls, and depleted in carbonyls (Jinich et al 2020).  Yet despite avoiding carbonyls, biosynthesis requires their reactivity. Most biosynthesis reactions involve a nucleophilic carbon attacking an (electrophilic) carbonyl.  This is the carbonyl paradox

L tracks the two-electron transfers from reductants. But the formal oxidation state (z) also tracks changes in the carbon skeleton. At a fixed L, every C-C coupling increments z and every C-C cleavage decrements it. Defining the total C-C bond order (n), one arrives at the helpful formula z = 2L – 4 + n. Formal oxidation state ranges from +4 in CO₂ to −4 in methane, and every electron added lowers it

These are carbon-level metrics. To describe a molecule, we can average the oxidation state of its constituents. Zc denotes the average formal oxidation state of a carbon molecule. Biomass sits in the middle. Bains & Seager (2012) scored ~4300 metabolites across a variety of organisms and found a narrow intermediate band, with the vast bulk of metabolites living near Zc = 0.0. Not every intermediate carbon is a metabolite. But biological intermediates are preferentially stable, with lower Gibbs energy (Jinich et al 2020). 
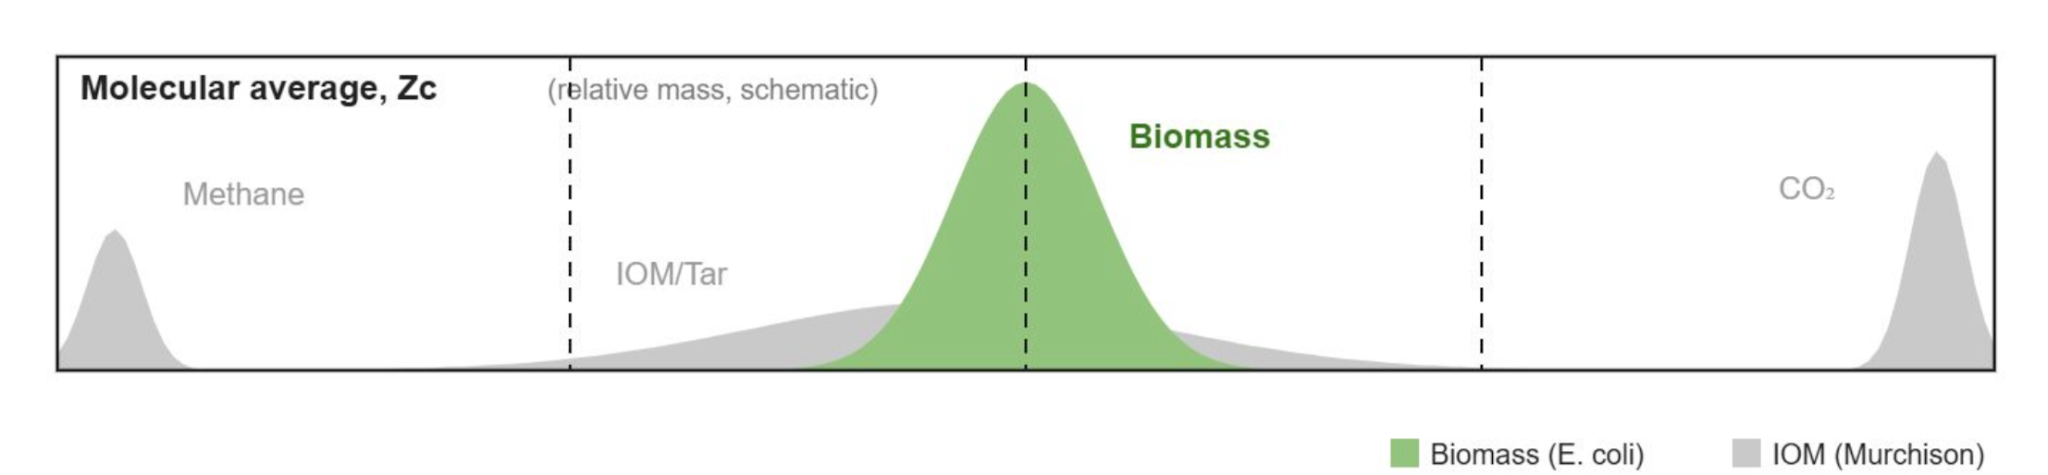
Biomass is deposited as L = 1 or 3. Biomass polarization explains how this is reconciled with Zc = 0. Fatty acids, amino acids, and anabolic carboxylates all have a reduced body around 0 and an oxidized tip at L = 3, with almost no carbon at L = 2. Sugars are the exception.

Why does life build intermediate carbon? It has tremendous combinatorial depth. Bains & Seager (2012) showed that, out of 1.7 million small molecules built from CHNOPS atoms, 95% were at intermediate oxidation state. This matters because useful biochemicals must satisfy many constraints at once. The solution space is sparse. Combinatorial depth makes selection possible. 

Intermediate carbon is also stable enough to persist in water. Water is gentle enough that most of carbon’s middle can persist. C–C, C–H, and many C–O bonds can survive long enough for complex molecules to accumulate and undergo selection (Pace 2001). Biomass polarization also helps. At cellular pH a carboxylate carries a negative charge, which keeps it dissolved in water and unable to leak through a membrane. Finally, Jinich et al. (2020) find metabolites more soluble than the nonbiological compounds of the same size that life skips.

Before carbon dioxide can be reduced, it must be activated. Activation has two components:

  1. Kinetic lability lowers the barrier to reaction. Lower barriers often dramatically speed up the rate of reaction.
  2. Thermodynamic driving makes a reaction energetically favorable.

Lability and driving are often correlated, but not always. Kinetic trapping describes a metastable molecule. These products “want” to react thermodynamically, but the activation barrier precludes the reaction at reasonable time frames. Nearly all biomass is kinetically trapped. For example, ATP hydrolysis is strongly favorable, yet it persists in water because the uncatalyzed reaction is slow. Biomass remains loaded, and can be harvested for energy once an enzyme opens the barrier.  

Driving does not require a change in oxidation state. A carboxylate is stable, but replacing its oxygen with a better leaving group produces an activated acyl compound such as a thioester or acyl phosphate. The carbon stays at L = 3 throughout.

In metabolism, intermediate carbon passes through the carbonyl hazard zone. It is quickly deactivated in a chemically selective manner. Without that control, abiotic carbon can go awry. In an aldol reaction, two carbonyls enter and one exits, so the reaction can continue indefinitely.  The chemistry fans outward. Dehydration gradually pulls the survivors into aromatic rings. The resultant products include tar or insoluble organic matter (IOM). Tar is a dead end. It can burn, but no biological reaction can return it to a set of small molecules. 

To convert carbon dioxide to biomass, or vice versa, carbon must first become labile. Anabolism requires three ingredients: a carbon source, an energy currency to activate it, and a reductant currency to attach electrons to it. Consider the Calvin cycle. NADPH is too weak a reductant for a carboxylate, so ATP first converts the carboxylation product, 3-phosphoglycerate, into 1,3-bisphosphoglycerate, which NADPH can reduce. Activation comes first; reduction follows; and the product, glyceraldehyde 3-phosphate, is an aldehyde inside the hazard zone. From there the carbon descends by condensation into the forms we know as biomass.

Catabolism does not simply oxidize biomass either. It must first activate the carbon, by phosphorylation or by attachment to coenzyme A, before the oxidations run. The descent returns far more ATP than the climb cost, and the electrons removed reload the reductant pool. Anabolism and catabolism trace the same loop in opposite directions.
Reductant Carriers

We are ready to return to electrochemistry. Galvanic cells convert redox disequilibria into a stock of ATP. Cells use this energy to fix carbon. 

The requirement to reduce carbon is in tension with the modularity of respiration. What if the respiratory substrate is not strong enough to reduce carbon?  H₂S sits at −0.27 V, NH₄⁺ at +0.34 V, NO₂⁻ at +0.43 V, and Fe²⁺ at +0.77 V in the acidic water where the best-studied iron oxidizers grow. All of these are far weaker donors than biosynthesis requires. How do these cells reduce carbon uphill?

To address this, life routes biosynthetic electrons through two universal reductant carriers. The nicotinamide cofactors NAD(P)H sit at −0.32 V under standard conditions. Ferredoxin, a small iron-sulfur protein, sits between −0.40 and −0.50 V depending on the organism. Once carbon is activated, NAD(P)H can reduce it, and it serves nearly all biosynthetic reductions. Ferredoxin covers the few steps that need a stronger reductant. After donating electrons to biosynthesis, these carriers return in their oxidized forms and must be reduced again.

A reduction carrier provides other benefits beyond enabling the reduction of carbon. The diversity of respiration enables flexible energy capture. But it would be inefficient to tune thousands of metabolic pathways to many dozens of environmental reductants. The reduction currency provides modularity: an adaptor layer between energy capture and biosynthesis. It also provides regulation. Rather than leaving reductants at the potential and concentration imposed by the local habitat, the cell can maintain its reductant pools at controlled redox states. For example, NAD(P)H potential is ideally suited to keep dangerous carbonyls at low steady-state concentration (Jinich et al 2018).

But how does a cell finance carrier recycling? When the environmental reductant is lower potential than the reductant carriers, carrier recycling doesn’t require energy. Hydrogen-eating autotrophs can produce NADH directly, because hydrogen at nominal conditions sits above NAD on the tower. Heterotrophs can reduce their carriers for free as well. The electrons in organic carbon span a wide range of potentials, and metabolism harvests only the deepest ones. 

But what happens when the environmental reductant is weaker than the carrier? In that situation, carrier recycling requires work. This is where life can manifest its metabolism as an electrolytic cell. 

The Electrolytic Cell

We met electrolytic cells in industrial reduction of alumina to aluminum. If stronger reductants are unavailable, the smelter can deploy external work, and leave that work embodied in the metal. If no stronger reductants are available to life, it can harness PMF from its galvanic cell to perform carrier recycling. Then biosynthesis drives CO₂ to biomass from the reductant carriers. Biomass is congealed electricity. 

When reductant recycling is not free, biology has found two ways to fund it with redox disequilibria.

First, reverse electron transport (RET) can pay from the wire. The PMF can push electrons from the quinone pool to NAD⁺, or from H₂ to ferredoxin. Complex I does this in nitrite and iron oxidizers. Ech does this in methanogens that carry cytochromes.

Second, electron bifurcation also taps energy metabolism, but pays earlier. Bifurcation takes an electron pair from a weak reductant and splits it: one electron falls to a high-potential acceptor, and the energy released pays for lifting the other onto ferredoxin.

Both of these solutions involve a redox disequilibrium powering two loads in parallel: synthesizing energy currency and reductant carriers. Energy metabolism and carbon metabolism compete. But there is another way to synthesize reductants without drawing on a redox disequilibrium. 

Light can serve the same purpose. A photosynthetic reaction center absorbs a photon and returns the electron at a lower potential than it received. The photon raises the electron by more than one volt, but part of that energy is spent to make the reaction irreversible. The resultant voltage gain (about 0.5 volts) subsidizes carrier recycling.  A chemotroph needs an environmental reductant above its carriers on the tower, or it pays for the lift with bifurcation or RET. But an anoxygenic phototroph can use an environmental reductant that is 0.5 volts weaker, because light pays for that part of the lift. 

Two families of reaction centers exist. 

Purple bacteria carry the type II reaction center, which deposits its electron in the quinone pool. The electron travels through the ETC cyclically, and produces PMF. Quinol is a weaker reductant than NADH, so reductant recycling must use RET. The purple bacterium converts PMF into reductant.

Green sulfur bacteria carry the type I reaction center, which deposits its electron in ferredoxin. Light pays for reductant recycling directly. Electrons reach the reaction center through the ETC, making PMF along the way. They come from the environmental reductant, or from ferredoxin sent back around the loop. The green sulfur bacterium spends reductant to make PMF, the reverse of the purple bacterium. 

Light arrives by day, but energy expenditure is continuous. Every phototroph relies on carbon-based storage to bridge the gap. Bacteria use glycogen; plants use starch. The stock is filled by electrolytic anabolism by day, and drained by galvanic catabolism by night. The structure mirrors the solar-power battery bank: two direct couplings separated in time by a stock. 

Oxygenic photosynthesis wired both reaction centers in series. The type II center moves electrons from water to the quinone pool. Their descent through the ETC produces energy. The type I center lifts the electrons into a ferredoxin carrier. These photons together pay the whole 1.25 volt span. This was the second bioenergetic revolution.
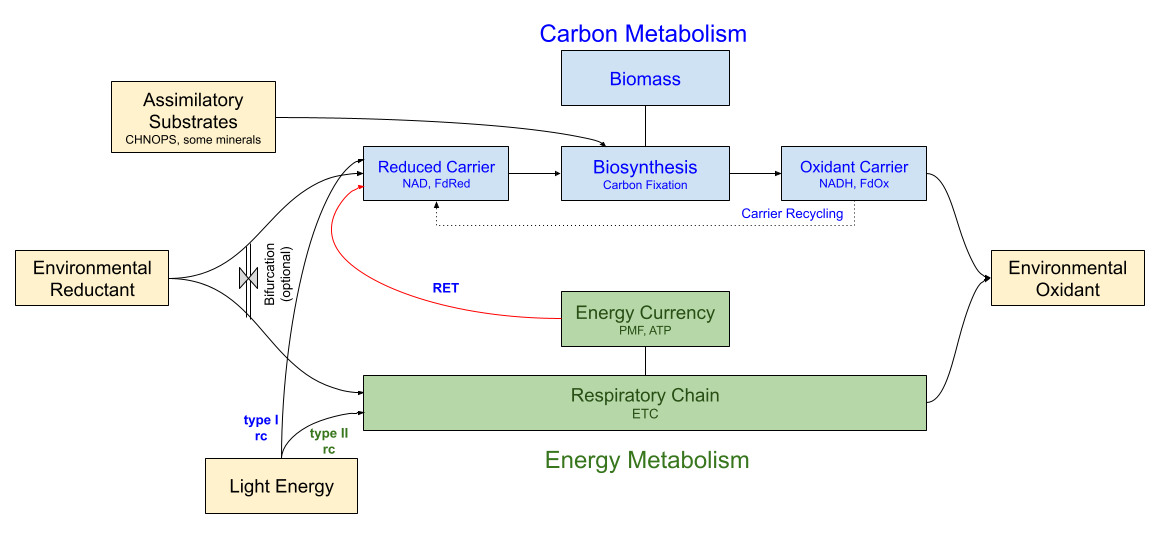
To sum up, galvanic and electrolytic circuits can coexist. The galvanic circuit harvests the environmental redox gradient as PMF. The electrolytic circuit uses PMF to lift electrons onto the carriers, to enable biosynthesis. A nitrite oxidizer runs both circuits continuously. A phototroph decouples galvanic and electrolytic circuits, with carbon-based storage as the “battery” stock in between.

One question remains. If life harvests redox gradients, what creates the gradients in the first place? 

A Planet-Sized Battery

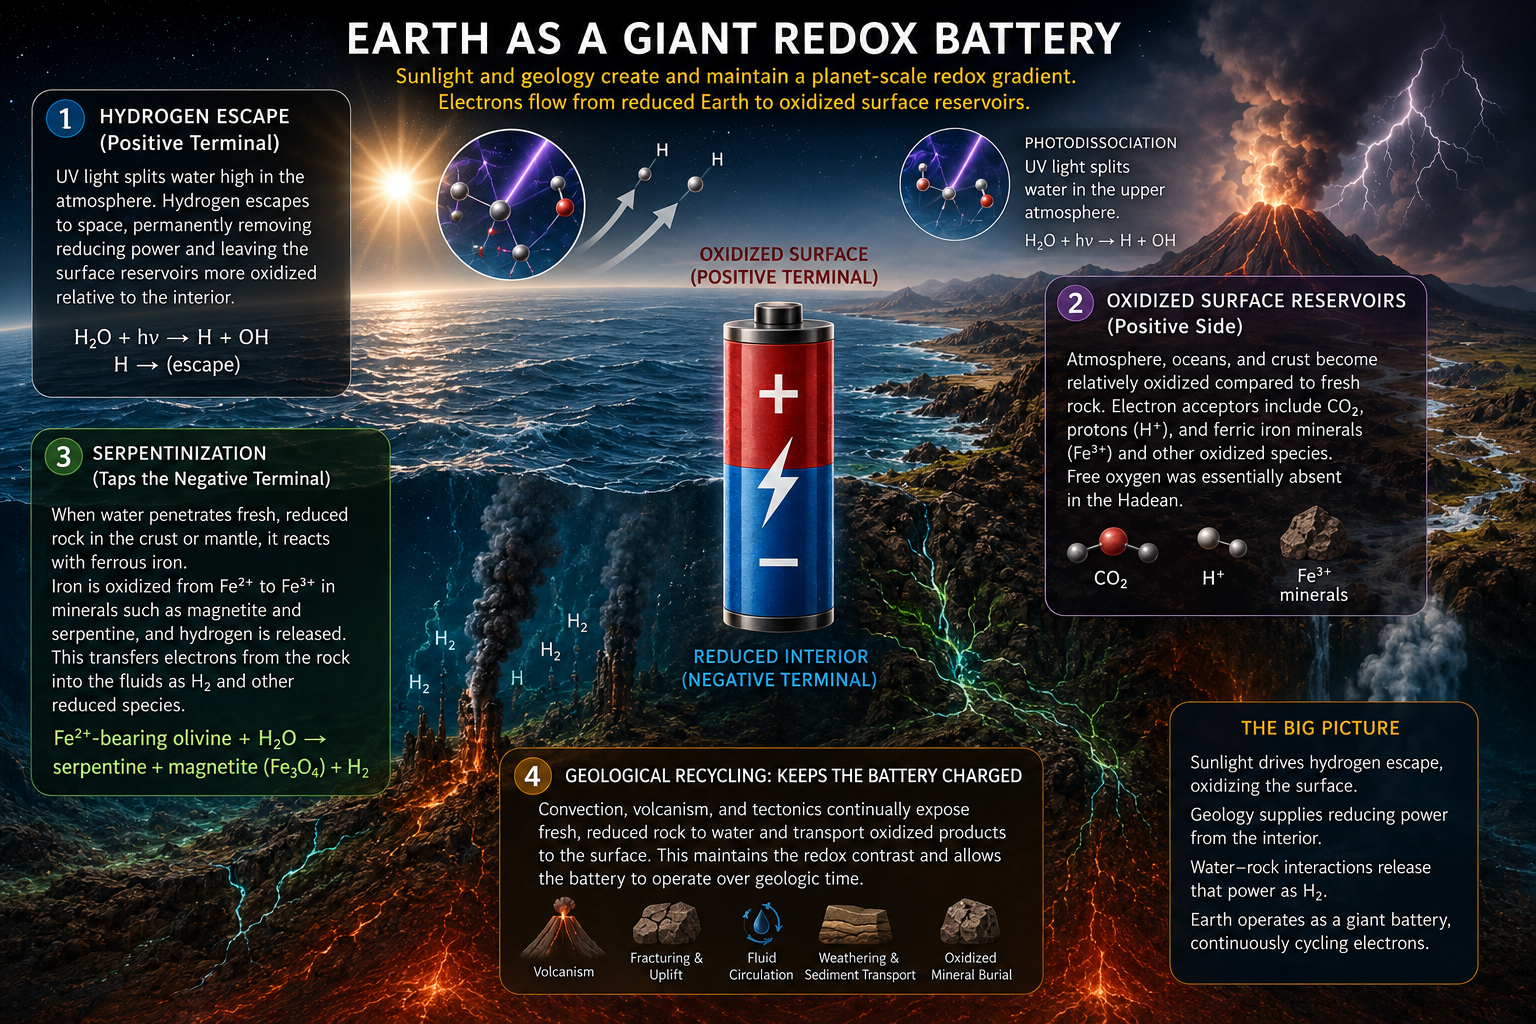
The battery existed before life learned to tap it. The Earth is a planet-sized redox battery (Smith & Morowitz 2016). Its anode is the mantle, rich in ferrous iron. Its cathode is the ocean and atmosphere, rich in carbon dioxide. The resultant redox span from H₂ to CO₂ is 0.2 volts. LUCA tapped this battery first. Plug-and-play respiration came later.

The atmosphere was birthed from volcanic degassing. This process built an oxidizing atmosphere of H₂O, CO₂, N₂, and SO₂. Photodissociation further oxidized the planetary cathode. Ultraviolet light breaks down water in the stratosphere. Hydrogen is prone to escape into the exosphere. The remaining oxygen is electron-hungry, oxidizing whatever it meets. Escaping hydrogen oxidizes our surface (Catling et al 2001). On Earth the process is slow, because the tropopause cold trap freezes water out before it reaches the stratosphere (Catling & Kasting 2017). Venus had no such trap, and photodissociation cost it an ocean (Kasting & Pollack 1983).

The mantle is rich with ultramafic rock, with a great deal of ferrous iron in olivine and pyroxene. Ferrous iron (Fe²⁺) holds one electron more than ferric iron (Fe³⁺), and loves to shed electrons. Serpentinization occurs when seawater reaches this rock. Ferrous iron gives its electron to water, ferric iron is left behind in magnetite and serpentine, and the electron leaves as H₂. H₂ carries the mantle’s reducing power, bringing the planetary anode up into the ocean. 

The two terminals do not discharge spontaneously. 4H₂ + CO₂ → CH₄ + 2H₂O is kinetically trapped. The reaction needs a catalyst. Life bridges the gap.

Other planets have iron and water, and some have freed oxygen by photodissociation. But corrosion usually stops. As the surface rusts, the electron-rich iron below becomes blocked. Current persists where fresh rock continues to meet water. On Earth, plate tectonics exposes fresh ultramafic rock at ridges, and subducts oxidized crust. But tectonics is not a hard requirement. Enceladus produces redox disequilibria from a porous core repeatedly damaged from tidal forces (Waite et al. 2017). 

Modern serpentinization yields ~10¹² mol of H₂ per year (Cannat et al. 2010; Worman et al. 2016). At 0.2 volts that is ~1.2 GW of chemical power. This planetary battery may have generated enough power to spark the first bioenergetic revolution (Russell et al. 2010).
Until next time. 

References

  • Bains & Seager (2012). A combinatorial approach to biochemical space: description and application to the redox distribution of metabolism.
  • Broda (1977). Two kinds of lithotrophs missing in nature.
  • Cannat et al (2010). Serpentinization and associated hydrogen and methane fluxes at slow spreading ridges.
  • Catling & Kasting (2017). Atmospheric Evolution on Inhabited and Lifeless Worlds.
  • Catling et al (2001). Biogenic methane, hydrogen escape, and the irreversible oxidation of early Earth.
  • Chance & Williams (1955). Respiratory enzymes in oxidative phosphorylation. III. The steady state.
  • Crockford et al (2023). The geologic history of primary productivity.
  • Falkowski et al (2008). The microbial engines that drive Earth’s biogeochemical cycles.
  • Froelich et al (1979). Early oxidation of organic matter in pelagic sediments of the eastern equatorial Atlantic: suboxic diagenesis.
  • Jelen et al (2016). The role of microbial electron transfer in the coevolution of the biosphere and geosphere.
  • Jinich et al (2018). Quantum chemistry reveals thermodynamic principles of redox biochemistry.
  • Jinich et al (2020). A thermodynamic atlas of carbon redox chemical space.
  • Kasting & Pollack (1983). Loss of water from Venus. I. Hydrodynamic escape of hydrogen.
  • Korenaga (2013). Initiation and evolution of plate tectonics on Earth: theories and observations.
  • Lane (2005). Power, Sex, Suicide: Mitochondria and the Meaning of Life.
  • Mitchell (1961). Coupling of phosphorylation to electron and hydrogen transfer by a chemi-osmotic type of mechanism.
  • Pace (2001). The universal nature of biochemistry.
  • Page et al (1999). Natural engineering principles of electron tunnelling in biological oxidation–reduction.
  • Russell & Hall (1997). The emergence of life from iron monosulphide bubbles at a submarine hydrothermal redox and pH front.
  • Russell et al (2010). Serpentinization as a source of energy at the origin of life.
  • Sleep et al (2004). H₂-rich fluids from serpentinization: geochemical and biotic implications.
  • Smith & Morowitz (2016). The Origin and Nature of Life on Earth: The Emergence of the Fourth Geosphere.
  • Strous et al (1999). Missing lithotroph identified as new planctomycete.
  • Thauer et al (1977). Energy conservation in chemotrophic anaerobic bacteria.
  • Waite et al (2017). Cassini finds molecular hydrogen in the Enceladus plume: evidence for hydrothermal processes.
  • Worman et al (2016). Global rate and distribution of H₂ gas produced by serpentinization within oceanic lithosphere.

Life’s First Ratchet

Part Of: Biology sequence
Followup To: A Portrait of LUCA
Content Summary: 4000 words, 20 min read

Cutting the Knot

Abiotic chemistry is leaky. Sugars brown into tar. Nucleic acid hydrolyzes in warm water on a timescale of days to weeks. Practically anything soluble diffuses away. Whatever complexity chemistry builds is temporary. To achieve LUCA, chemistry needed a ratchet.

The natural place to look for that ratchet is LUCA’s own subsystems, but they depend on one another. Genes prescribe proteins, yet cannot be expressed without proteins to transcribe and repair them. Metabolism supplies the materials for both, yet runs on protein catalysts that genes encode. Asking which came first is really asking which subsystem supplied the first ratchet. 

This Gordian knot is a question of functional priority. There are two ways to cut it.

One answer sees the first ratchet in nucleic acid. A system that can copy information can preserve innovation. This is the genetics-first tradition.

The other sees it in metabolism. A network that captures free energy can maintain its own organization and generate the building blocks heredity would later need. This is the metabolism-first tradition. 

The RNA World

Let’s set metabolism aside for a moment. Among the three biopolymers of the central dogma – DNA, RNA, and modern coded proteins – which came first? On this smaller question, the scientific community has achieved consensus. Metabolism-first and genetic-first camps agree: RNA came first. 

RNA is the answer because it can do both jobs. RNA stores information in its nucleotide sequence, like DNA. But it also folds itself into structures that catalyze chemical reactions, like proteins. The discovery of self-splicing introns revealed that RNA can act as a catalyst (Kruger et al. 1982). Most modern biological catalysts are protein enzymes. RNA catalysts are called ribozymes.  The universal coenzymes hint at the same history. NAD, FAD, coenzyme A, and SAM are all built on nucleotide scaffolds, which White (1976) read as molecular fossils of an RNA-based metabolism.

The ribosome is the engine of the genetic code. Remarkably, we are able to reconstruct its development at the atomic resolution. The oldest part of the ribosome, its center known as the peptidyl transferase center (PTC), contains no proteins at all. Its catalytic heart is entirely RNA. The ribosome is a ribozyme (Nissen et al. 2000). 

Other genetic polymers could, in principle, have preceded RNA. One genetic system could also have replaced another while preserving the basic function of heredity, a process sometimes called genetic takeover (Hud et al. 2013). But for now, in the name of parsimony, we will assume the first genetic ratchet was RNA. 

Why is protein-first no longer an active research tradition? Amino acids and short peptides may well predate RNA. But RNA apparently precedes modern coded proteins. The RNA World constitutes major progress in cutting the knot. The RNA world is silent on the order of metabolism. This is the residue. 

The genetics-first solution advocates a strong version of the RNA World: genetic heredity precedes the emergence of metabolism. Natural selection then improves the chemical system around the replicator. No one questions the capacity of nucleic acids to build and retain complexity. The burden is to show how geochemistry alone can birth such a wondrous digital inheritance

The metabolism-first tradition holds that metabolism existed before RNA, and therefore before genetically encoded proteins. A non-enzymatic protometabolism must exist first. This school fully accepts digital inheritance. But it proposes that protometabolism served as the first ratchet. Geochemistry is thus less burdened. The burden here is twofold: to show that non-enzymatic protometabolism is viable, and to show that a mere reaction network can preserve innovation without genes. We take up the second question in a later post.

Let us explore the viability of specific genetics-first and metabolism-first theories. These are grounded in specific habitats. But before we search for the Geography of Genesis, we must first understand what success looks like. Designing such a rubric takes us back to first principles. We begin with thermodynamics.

Life as a Dissipative Structure

Free energy is the part of energy that can do work. The second law of thermodynamics says that energy tends to spread out. Hot objects cool, gases mix, and chemical gradients become weak. 

A closed system does not exchange matter with its surroundings. Without a continuing source of usable energy, its chemical differences decay toward equilibrium. As a system approaches equilibrium, free energy is dissipated, often as heat, and less remains available to do useful work. 

Life manifests tremendous complexity and order. In What Is Life?, Erwin Schrödinger explained that life is not a closed system. An open system can exchange energy and matter, so an external process can maintain nonequilibrium conditions. Life exists in an open system.  It consumes external energy and matter, then releases heat and waste. Life keeps local order while disorder increases in its surroundings. 

The chemist Ilya Prigogine developed the idea of dissipative structures. These structures import free energy from their environment. They maintain local order by using this energy and releasing heat and waste into their surroundings. When the flow stops, the structure usually weakens or disappears. 

Dissipative structures occur at many scales. Examples include convection cells, flames, chemical oscillations, hurricanes, and living organisms. For the origin environments we will consider, two energy sources matter most: sunlight and geological activity. 

Our first requirement is sustained free-energy flux. The Garden of Eden must feature continuous (or at least recurrent) throughput sufficient to maintain far-from-equilibrium chemistry.

Chemical Selectivity

Free energy flux can drive useful reactions. It can also drive thousands of useless ones. 

In organic chemistry, a small set of reactive molecules can combine in many different ways. Each new product can then react with the original molecules or with other products. The number of possible pathways can grow rapidly. Researchers call this a combinatorial explosion

Scientific experiments (e.g., Miller Urey) and natural phenomena (e.g., carbonaceous chondrites) both tend to produce tar: complex mixtures that are difficult to separate or even characterize. 

The formose reaction gives another example. Recall that ribonucleic acid (RNA) uses one particular sugar: ribose. Under alkaline conditions, formaldehyde produces dozens of different sugars and related compounds, while ribose appears as only one component of the diverse products. As the reaction continues, useful products can disappear into still more complex chemistry. 

Life is different. Biology uses an extraordinarily restricted region of chemical space. About 500 naturally occurring amino acids have been identified, but the standard genetic code uses only 20 of these to build proteins. The same organizing principle appears in metabolism. Smith and Morowitz (2004) identify eleven TCA-cycle carboxylic acids as a universal anabolic core, from which the major pathways of biosynthesis branch. In their formulation, lipids, sugars, amino acids, nucleotides, and other major classes of biomolecules are ultimately built outward from this small conserved core. 

A biochemical environment must channel significant flux toward its target products, while limiting competing reactions. This is a second requirement for Eden: chemical selectivity

For genetics-first, this requirement must be met before selection can start. A nucleic acid is a monument of chemical selectivity: one sugar among dozens, a few bases among many, the correct linkage among several. The environment must achieve that discrimination unaided. Genetics-first chemistry therefore requires prebiotic mechanisms that narrow the product distributions. 

Metabolism-first theories propose another possibility. Positive feedback reactions (aka autocatalysis) can deliver selectivity without a special environment. But it only works if the network reinforces a restricted set of productive reactions, rather than amplifying chemical clutter. We will explore this topic in more detail in a followup post.

Deriving the Rubric

We can say more. Chemical principles allow us to derive five additional requirements. 

  1. Provenance. The required feedstocks and catalysts must arise or arrive naturally.
  2. Reactant Concentration. Feedstocks must reach concentrations high enough for useful reactions.
  3. Material Flux. Fresh material and free energy must continue to enter, while waste leaves and spent material is removed, reactivated, or recycled. 
  4. Product Retention. Useful products must also remain close by long enough to interact again, rather than immediately dispersing into the environment. 
  5. Net Production. Eden must not only produce these target products. It must produce them faster than it destroys them. We require net production.

Material flux and product retention are in creative tension. Eden must eject waste while selectively retaining useful products. 

The above concerns single reactions. But theories of abiogenesis rely on many processes.

Suppose one reaction requires acidic conditions, another alkaline conditions. Or one may require intense ultraviolet radiation, while another important product is rapidly destroyed by the same radiation. A theory may need to confine different reactions to different microenvironments. Such conflicts do not make a pathway impossible, but they do require that these microenvironments exchange products in a realistic way. This is the requirement for condition compatibility.

It is not enough to demonstrate each reaction separately in a laboratory. Products of one step must naturally reach the next without artificial purification, isolation, or reagent replacement.  This is the requirement for process continuity.

A favorable environment is of little use if it disappears before cumulative organization can develop. The environment must persist or recur often enough to provide continuity across time. Our last requirement is habitat persistence

These requirements are universal. But genetics-first and metabolism-first theories also generate idiosyncratic requirements. Let’s examine each in turn.

Building Nucleic Acid

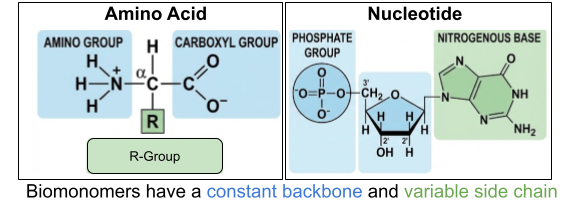
Genetics-first demands a specific product. Geochemistry must synthesize a nucleotide. Every nucleotide brings together a sugar (ribose), a nitrogenous base, and a phosphate group into a single large molecule:

Carbon, hydrogen, and oxygen are readily available in many geological habitats. Nucleotides also require nitrogen and phosphorus, and these present new challenges for genetics-first models.

Most environmental nitrogen is molecular nitrogen, N₂. The strong triple bond in N₂ makes it unusually unreactive. Before nucleotide synthesis can begin, some process must supply fixed nitrogen in a reactive form. Nitrogen fixation is one way to make it more reactive (Mateo-Marti et al. 2019). Similarly, phosphorus is present in rocks, but much of it occurs in minerals with low solubility. In water containing calcium, dissolved phosphate is quickly locked away into minerals. Many prebiotic reactions, meanwhile, require phosphate concentrations far above those found in ordinary natural waters. This is the phosphate problem (Toner and Catling 2020). Together, these problems generate two requirements. A genetics-first theory requires bioavailable nitrogen and phosphate. 

Of course, individual nucleotides aren’t enough. They must come together as long strands of nucleic acid

RNA polymerization occurs through a condensation reaction. But condensation polymerization is disfavored in water. The equilibrium instead favors the reverse reaction, hydrolysis. Joining nucleotides into a chain therefore requires an input of free energy (Yadav et al. 2020). Monomer activation denotes the requirement to store or couple this free energy so that phosphodiester-bond formation can proceed.

Providing free energy is not enough. Polymerization also requires high efficiency. A 45-nucleotide polymer requires 44 bond-forming steps. At each step, the chain can degrade. Even modest inefficiency therefore compounds across the chain. Polymer elongation denotes the requirement for bond formation to be efficient enough to produce functional-length polymers.

Towards a Replicator

We’ve established a long RNA polymer. Next, it must copy itself. 

Natural selection cannot act on sequence information until polymers are long enough for different sequences to produce different functional structures. Very short oligomers have limited capacity to fold into complex three-dimensional forms.

We do not know a strict minimum length for a useful hereditary polymer. But Gianni et al. (2026) provide an important empirical benchmark. They discovered QT45, a polymerase ribozyme only 45 nucleotides long. QT45 can catalyze synthesis of both its own sequence and its complementary strand.

Recall that nucleotides attach to their complements via Watson-Crick pair bonds. These bound duplexes are very stable. This stability assists during the replication, but presents a problem afterwards. If the strands remain bound, neither is available as a template for the next generation. This is the strand inhibition problem. For replication to persist, the resulting duplex must separate without damaging the sequence. Template reset denotes this requirement. 

A catalyst that helps every molecule around it, freeloaders included, gains nothing from its own innovation. Some mechanism must couple products to their producers: what a system makes must preferentially benefit that system. Compartments are the obvious mechanism, but mineral pores, surfaces, and reaction-diffusion gradients are candidates too. As we will see in a later post, this requirement bites both camps, and selection will demand a sharper version of it.

So ends our discussion of genetics-first. Now we turn to metabolism-first theories.

The Twin Engines of Protometabolism

Genetics-first requires a specific deliverable: self-replicating RNA. Where is the analogue for metabolism-first? Which seeds accreted the fantastic complexity of modern metabolism?

Metabolic networks are not tangled webs. Their architecture is hierarchically modular: tightly linked modules nest within larger modules, all organized around a small connective core (Ravasz et al. 2002). Architectures like this arise when networks expand by attachment, each new pathway bolting onto machinery that already exists. And once a dozen downstream pathways depend on an upstream reaction, that reaction is frozen in place; changing it breaks everything built on top. Wimsatt (2007) calls this generative entrenchment. Entrenchment gives the network a readable stratigraphy. Deeply embedded core chemistry should tend to be older than dependent peripheral pathways. We will meet the same logic again when we explore the ribosome’s history from its onion-like layers. 

All living things rely on energy metabolism to fuel themselves and anabolism to build up complex organics. The differences reside where the carbon comes from. Autotrophs use carbon fixation to build biomass from inorganic carbon such as CO2. Heterotrophs obtain reduced carbon from the environment, and rely on catabolism to break down the complex organics in their diet. 

The modern biosphere rests on an autotrophic foundation. Nearly all carbon enters through the fixation of CO2. Life does not depend significantly on extraterrestrial, atmospherically or geologically produced organics. But on the Hadean Earth, impact delivery supplied organics at a far higher rate. Oparin and Haldane’s original vision of primordial soup presumed a heterotrophic origin. So which was it? Did the first life eat chondrites or fix carbon? 

Two converging lines of evidence suggest fixation. The first comes from genes and energetics. Weiss et al. (2016) traced 355 gene families back to LUCA and recovered the profile of an anaerobic, H2-dependent autotroph. Phylogenetic reconstructions are often contested. But Wimmer et al. (2021) computed the thermodynamics of LUCA’s ~400 core reactions and found that roughly 96% run exergonically under H2-rich anoxic conditions. The genes describe an autotroph, and the metabolism they encode fits an autotroph’s environment.

The second line of evidence applies our stratigraphy. Catabolism sits at the periphery. Major catabolic pathways characteristically converge on the universal anabolic core. Moreover, the substrates it digests are sugars, lipids, and proteins: the characteristic products of anabolism (Schönheit, Buckel & Martin 2016). The enzyme record concurs. The ancestral fructose-bisphosphate enzyme of archaea is locked into the gluconeogenic direction, marking glycolysis as a later retrofit (Say & Fuchs 2010). 

Taken together, these suggest that anabolism came first. The first life was likely autotrophic.

Carbon fixation places specific requirements on the environment. It requires both CO2 and a continuous electron supply. Candidate electron donors (reductants) include H2, Fe(II), and H2S. Reductant and carbon co-delivery denotes the requirement that electron donors and inorganic carbon arrive together, continuously, at rates sufficient to sustain fixation.

Anabolic metabolism synthesizes thousands of biological molecules. The stratigraphy points inward: the oldest chemistry of all should sit at the very center of the network. Smith and Morowitz (2004) identify it. Eleven carboxylic acids form a universal anabolic core.

With a handful of glycine-related exceptions, all biosynthesis radiates from five of these acids. Acetate seeds the lipids; pyruvate the sugars and the alanine-family amino acids; oxaloacetate the aspartate family, and through aspartate the pyrimidines; α-ketoglutarate the glutamate family; succinyl-CoA the pyrroles. (Purines are the untidy exception, assembled from glycine, aspartate, glutamine, and the same C1 chemistry owned by the WL pathway) These are the pillars of anabolic metabolism

The entire biosphere uses seven carbon fixation pathways (CBB, rTCA, WL, 3-HP, 3-HP/4-HB, DC/4-HB, and roTCA). Which of these was used by LUCA? 

You may recall learning about the tricarboxylic acid (TCA) cycle (aka Krebs cycle) at school. In modern organisms, this cycle is catabolic. For decades that was its only known direction. But Evans et al (1966) and Shiba et al (1985) discovered it also runs in the reverse direction. This rTCA cycle is an autotrophic engine of carbon fixation. The eleven carboxylic acids are precisely the universal anabolic core. It is a strong candidate for the seed of protometabolism. 

Several steps of any fixation pathway are endergonic (thermodynamically dysfavored) under ordinary conditions. Modern cells pay for these steps with ATP hydrolysis. A protometabolism has no ATP. The environment itself must pay. Thermodynamic drive denotes the requirement that environmental chemistry makes the overall pathway favorable without modern energy currencies like ATP.

We have identified the rTCA cycle by reading the network’s stratigraphy. Comparative biology provides another glimpse. Consider organisms that are anaerobic and chemoautotrophic. In archaea, these are the methanogens; in bacteria the acetogens. What do they have in common? 

Both methanogens and acetogens rely on the Wood-Ljungdahl (WL) pathway (aka acetyl-CoA pathway). WL is in fact the only carbon-fixation pathway found in both bacteria and archaea. Weiss et al (2016) place WL in LUCA itself. WL is also thermodynamically unique. Every other fixation pathway runs uphill and is financed with ATP. The Calvin cycle spends roughly nine ATP per pyruvate; rTCA is among the cheapest. WL runs downhill. Under anoxic conditions with H2 as the electron donor, the overall reaction is exergonic, releasing roughly 100 kJ/mol. Acetogens and methanogens exploit this: the same pathway that builds their biomass also charges their membranes and makes their ATP.  A cell running WL on a H2 supply occupies what Everett Shock called a “free lunch you are paid to eat.” WL is the only fixation chemistry that could have paid its own way. 

Finally, methanogens and acetogens also share a second piece of machinery: flavin-based electron bifurcation. The problem it solves is that reducing CO₂ requires electrons at potential lower than H₂ itself can supply. Bifurcation takes an electron pair from a mid-potential donor and splits it: one electron falls to a high-potential acceptor, and the energy released pays for lifting the other onto low-potential ferredoxin, the reductant that carbon fixation demands. Its distribution across both domains marks it as ancient (Buckel & Thauer 2013).

These two engines have complementary strengths and weaknesses. rTCA contains the entire anabolic core, and it is autocatalytic. But it runs uphill. WL has the right energetics: it is the only chemistry that runs downhill. But it is a linear, non-autocatalytic path, and hence cannot auto-amplify. Braakman and Smith (2013) place a fused WL-rTCA network at the base of the tree. For now, we will treat these fixation pathways as complementary halves of one machine.

Both WL and rTCA are now executed by enzymes. But buried inside many of these proteins are metal-sulfur cofactors, and these are often the catalytically active sites. Ferredoxin contains cubanes whose geometry resembles motifs found in  the mineral greigite (via its precursor mackinawite). The nickel-iron-sulfur cluster at the heart of CO dehydrogenase and acetyl-CoA synthase likewise resembles violarite. Zhao et al. (2022) quantified these cofactor-mineral correspondences. These structural correspondences hint at evolutionary continuity. Russell and Martin (2004) argue that these catalytic sites are mineral fossils, and the surrounding protein is later packaging around an ancient inorganic core. 

Metabolism-first envisions a non-enzymatic protometabolism. Geochemical continuity suggests the earliest catalysts were mineral. But modern enzymes accelerate reactions by factors up to 10^17. Without them, most reactions proceed at negligible rates. Kinetic accessibility denotes the experimental requirement that mineral, metal, or small-molecule catalysts sit in the flow path and drive the key reactions at meaningful rates. 

Vigorous flow solves one problem and creates another. The same throughput that delivers reductant and exports waste can wash a network’s working parts away faster than the chemistry regenerates them. A pathway turns on a characteristic timescale, and its intermediates must survive long enough to complete the loop. Intermediate retention denotes the requirement that residence times exceed network turnover times.

The deepest layer of the network has another striking property. If we abstract away the modern enzymes, cofactors, and carrier groups that now mediate it, the carbon skeletons of WL and rTCA contain only carbon, hydrogen and oxygen. These skeletons are strictly a CHO metabolism. Nitrogen is not needed to construct this central carbon framework. It seems to have entered later, converting pre-existing carbon skeletons into amino acids and the broader CHON metabolism (Braakman & Smith 2013).  Phosphate may show a similar pattern: Goldford et al. (2017) reconstructed a plausible ancient metabolic network with little dependence on phosphate, although Tian et al. (2019) contest how far that inference can be pushed.  This is compatible with de Duve (1991)’s hypothesis that thioesters (like acetyl-CoA) served as an energy currency predecessor of ATP. 

The resulting picture is one of elemental expansion: a CHO carbon-skeleton core, followed by nitrogen incorporation and increasingly phosphate-dependent chemistry. This ordering is naturally compatible with metabolism-first. If RNA came first, despite already requiring fixed nitrogen and phosphate, it becomes less obvious why the deepest metabolic layers should preserve a simpler elemental architecture. 

A Scorecard For Habitat

We have generated twenty requirements.

Universal (11)

  • Sustained free-energy flux. Continuous or recurrent throughput maintaining far-from-equilibrium chemistry.
  • Chemical selectivity. Flux channeled toward targets, against the combinatorial tar.
  • Provenance. Feedstocks and catalysts arise or arrive naturally.
  • Reactant concentration. Feedstocks reach reaction-viable concentrations.
  • Material flux. Energy and material in, waste out, spent material recycled.
  • Product retention. Useful products stay long enough to react again (in creative tension with material flux).
  • Net production. Target made faster than destroyed.
  • Condition compatibility. Incompatible reactions confined to microenvironments that exchange products realistically.
  • Process continuity. Each chemical step connects in geochemically plausible way, without artificial laboratory techniques.
  • Habitat persistence. Eden either outlasts, or recurs frequently.
  • Localization. Products must preferentially benefit the systems that produce them, rather than diffusing to competitors. 

Genetics-first (5)

  • Bioavailable nitrogen. Reactive C–N precursors.
  • Bioavailable phosphate. Soluble, concentrated phosphate despite mineral lock-up.
  • Monomer activation. Free energy stored or coupled so phosphodiester bonds can form.
  • Polymer elongation. Bond formation is efficient and repeatable to functional lengths.
  • Template reset. Duplexes separate non-destructively for the next round.

Metabolism-first (4)

  • Reductant and carbon co-delivery. Electron donors and CO₂ arrive together, continuously.
  • Thermodynamic drive. The environment pays for endergonic chemistry without ATP.
  • Kinetic accessibility. Non-enzymatic catalysts must drive key reactions at meaningful rates.
  • Intermediate retention. Residence times exceed network turnover times.

Next post, we will use this scorecard to grade specific hypotheses from both camps. See you then!

References

  • Braakman & Smith (2013). The Compositional and Evolutionary Logic of Metabolism.
  • Buckel & Thauer (2013). Energy Conservation via Electron-Bifurcating Ferredoxin Reduction and Proton/Na⁺-Translocating Ferredoxin Oxidation.
  • Butlerov (1861). Formation Synthétique d’une Substance Sucrée.
  • de Duve (1991). Blueprint for a Cell: The Nature and Origin of Life.
  • Evans, Buchanan & Arnon (1966). A New Ferredoxin-Dependent Carbon Reduction Cycle in a Photosynthetic Bacterium.
  • Gianni et al. (2026). A Small Polymerase Ribozyme That Can Synthesize Itself and Its Complementary Strand.
  • Goldford, Hartman, Smith & Segrè (2017). Remnants of an Ancient Metabolism without Phosphate.
  • Hud et al. (2013). The Origin of RNA and “My Grandfather’s Axe.”
  • Kruger et al. (1982). Self-Splicing RNA: Autoexcision and Autocyclization of the Ribosomal RNA Intervening Sequence of Tetrahymena.
  • Mateo-Marti et al. (2019). Pyrite-Induced UV-Photocatalytic Abiotic Nitrogen Fixation: Implications for Early Atmospheres and Life.
  • Nissen et al. (2000). The Structural Basis of Ribosome Activity in Peptide Bond Synthesis.
  • Pitsch et al. (1995). Mineral Induced Formation of Sugar Phosphates.
  • Ravasz et al. (2002). Hierarchical Organization of Modularity in Metabolic Networks.
  • Russell & Martin (2004). The Rocky Roots of the Acetyl-CoA Pathway.
  • Say & Fuchs (2010). Fructose 1,6-Bisphosphate Aldolase/Phosphatase May Be an Ancestral Gluconeogenic Enzyme.
  • Schönheit, Buckel & Martin (2016). On the Origin of Heterotrophy.
  • Shiba et al. (1985). The CO₂ Assimilation via the Reductive Tricarboxylic Acid Cycle in an Obligately Autotrophic, Aerobic Hydrogen-Oxidizing Bacterium, Hydrogenobacter thermophilus.
  • Smith & Morowitz (2004). Universality in Intermediary Metabolism.
  • Tian et al. (2019). Phosphates as Energy Sources to Expand Metabolic Networks.
  • Toner & Catling (2020). A Carbonate-Rich Lake Solution to the Phosphate Problem of the Origin of Life.
  • Weiss et al. (2016). The Physiology and Habitat of the Last Universal Common Ancestor.
  • White (1976). Coenzymes as Fossils of an Earlier Metabolic State.
  • Wimmer et al. (2021). Energy at Origins: Favorable Thermodynamics of Biosynthetic Reactions in the Last Universal Common Ancestor.
  • Wimsatt (2007). Re-Engineering Philosophy for Limited Beings: Piecewise Approximations to Reality.
  • Yadav et al. (2020). Chemistry of Abiotic Nucleotide Synthesis.
  • Zhao et al. (2022). Quantifying Mineral-Ligand Structural Similarities: Bridging the Geological World of Minerals with the Biological World of Enzymes

A Portrait of LUCA

Part Of: Biology sequence
Content Summary: 3000 words, 15 min read

The Basic Facts of Abiogenesis

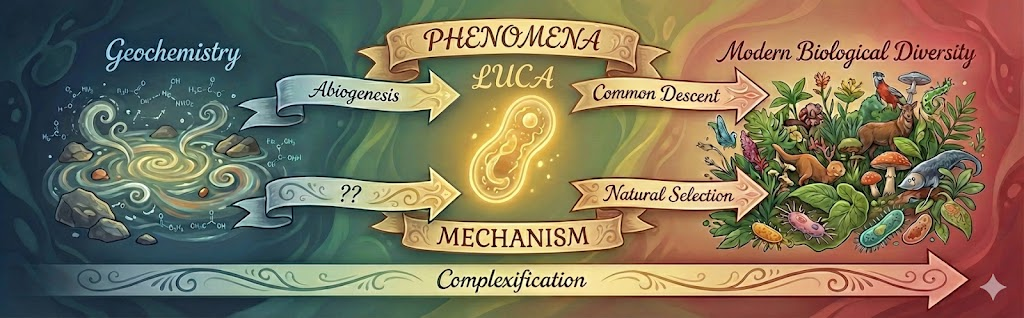
From the geochemistry of the early Earth, life emerged. The abiogenesis phenomenon is interesting in at least two contexts:

First, it is one of the most difficult unsolved scientific problems in the 21st century. Common descent describes how all of the marvelous diversity of biology stem from a single source: a single celled organism called the Last Universal Common Ancestor (LUCA). We have a clear mechanistic understanding for how this complexification was possible: evolution by natural selection

But natural selection relies on the machinery of genetic inheritance to produce complexity. How was it possible to create the sophisticated nanomachinery of LUCA (e.g., ATP synthase) before natural selection took effect? What other process besides natural selection can produce complexification?

Second, abiogenesis research is relevant to questions in astrobiology:

Could extraterrestrial life have begun much earlier? 

  • Gen 1 stars (born 13.5-13 Ga) had no exoplanets because the heavy elements required for planetary cores didn’t exist (supernova nucleosynthesis arrived later). 
  • Gen 2 stars (born 13-10 Ga) may have been able to support life, but these exoplanets were small, terrestrial, and less metallic – unable to furnish geomagnetism nor plate tectonics. 
  • Gen 3 stars (born 10-0 Ga) can support life. Exoplanets in this era now include gas giants, and are highly metallic. Geomagnetism protects us from UV radiation, and plate tectonics promote mineral diversity. 

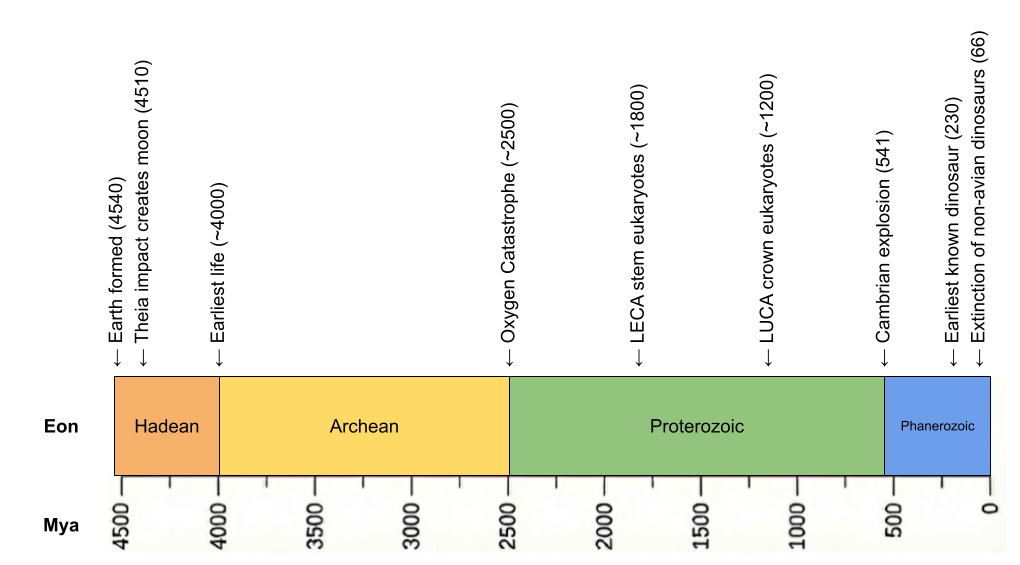
Is abiogenesis easy or hard? The Earth was born 4.6 Ga, with the Theia impactor at 4.54 Ga creating the moon and the Moneta impactor at 4.51 Ga providing siderophilic veneer. Despite such impacts, zircon evidence now suggests a cool early earth, with oceans appearing as early as 4.4 Ga. Critically, our evidence for life arrives very early (Isua banded-iron formations at 3.8 Ga, enriched carbon-12 in zircon graphite at 4.1 Ga). Abiogenesis occurred only ~500 million years after the sterilizing impact of Moneta! Speed is evidence of ease.

If the universe has billions of potentially habitable planets, why haven’t we found any aliens? One explanation to the Fermi Paradox is the Great Filter – some incredibly difficult barrier that prevents life from becoming spacefaring. The scary part is we don’t know where this filter is: behind us (we got incredibly lucky to make it this far) or ahead of us (something typically destroys civilizations before they can spread across the galaxy). If abiogenesis is indeed easy, this might mean the Great Filter is ahead of us.

LUCA from Paleobiogeography

To understand the journey of abiogenesis, we must first understand the destination. What do we know about LUCA?

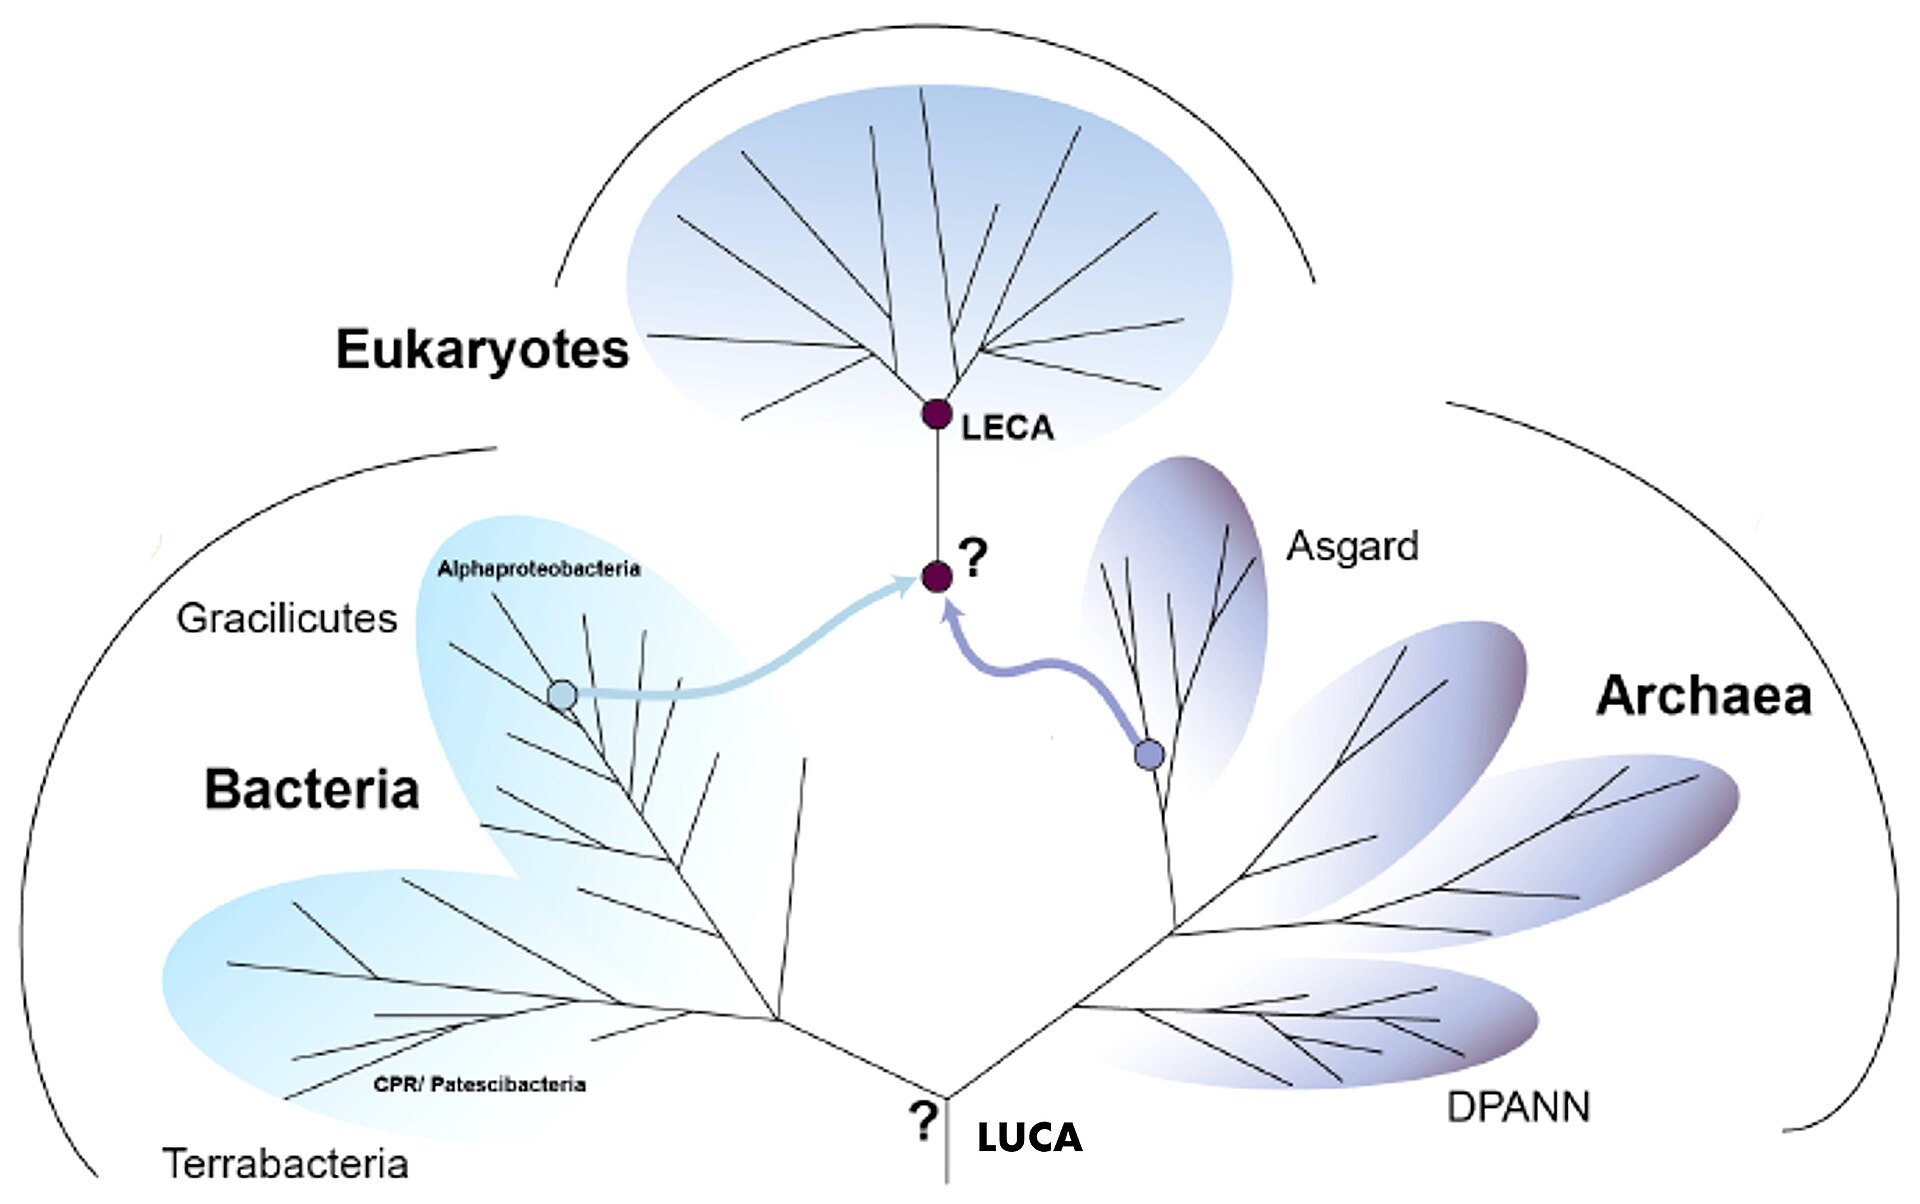
The history of life constrains our search. Two domains of simple life (prokaryotes) emerged in the Hadean: bacteria and archaea. But eukaryotes (complex life) emerged much later, as a result of endosymbiosis between the two families. 

Life was microbial for the first 80% of Earth’s history. But in 1.0 Ga, multicellularity was invented. Starting at that time, multicellularity was invented multiple times, but it only really took off in crown group eukaryotes. 

LUCA must have been anaerobic. For the first half of Earth’s history, oxygen was locked in water and rocks – there was no free oxygen in the atmosphere. Only when photosynthesis caused the Oxygen Catastrophe in 2.4 Ga did atmospheric oxygen go from 0 to 10%. 

LUCA is simple (prokaryotic), microbial (unicellular), and anaerobic.

LUCA from Phylogenetics

We can learn more about LUCA from genetic analyses. 

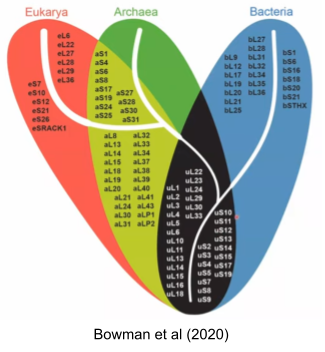
Genes conserved across all species constitute what is called the Universal Gene Set of Life (UGSL), and consists of less than 100 genes (Harris et al 2003). Not surprisingly, the UGSL is dominated by translation-related genes. Here are those ribosomal genes in black:

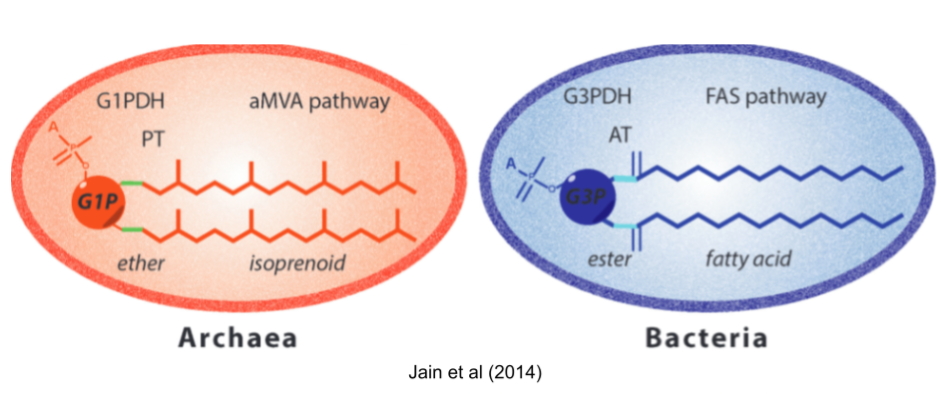
Many differences between archaea and bacteria arose after they diverged. But some differences are more troubling. First among these is the Lipid Divide. Both domains build cells with the phospholipid bilayer membranes, so it is natural to suspect that LUCA had a similar coat. But almost all biochemical details are mirrored! They use different glycerol backbones, different hydrophobic chains (isoprenoid vs fatty acids), different links to those chains (ether vs ester), and different biosynthetic pathways (FAS+AT vs aMVA+PT). 

There are three major theories for the lipid composition of LUCA:

  • Heterochiral theories. LUCA had both lipid biochemistries available. Heterochiral membranes were recently found to be viable (Caforio et al 2018). 
  • Thermoreduction theories. Many archaea are adapted to extreme environments. So LUCA had a bacterial phospholipid, and the more robust archaeal lipids were derived to support more extreme niches.
  • Protocompartment theories. LUCA didn’t use phospholipids at all! It used a simple fatty acid membrane, or coacervate droplets, or mineral pores to achieve compartmentalization.

Another divergence is even more troubling: bacteria and archaea use completely different DNA replicase proteins (Forterre et al 2013). 

Neither divide has a satisfying resolution. Each of the three lipid hypotheses faces serious objections, and the DNA replicase situation is arguably worse: there is no consensus account for how two non-homologous replication machineries could descend from a single ancestor, which has led some (e.g., Forterre 2006) to argue that DNA itself was a viral invention acquired independently by the bacterial and archaeal lineages. Any portrait of LUCA that papers over these divides is overconfident.

LUCA from Biochemistry

At the atomic level, organisms are built with CHNOPS: Carbon, Hydrogen, Nitrogen, Oxygen, Phosphorous, and Sulfur. 

At the molecular level, life is composed of three biopolymers.

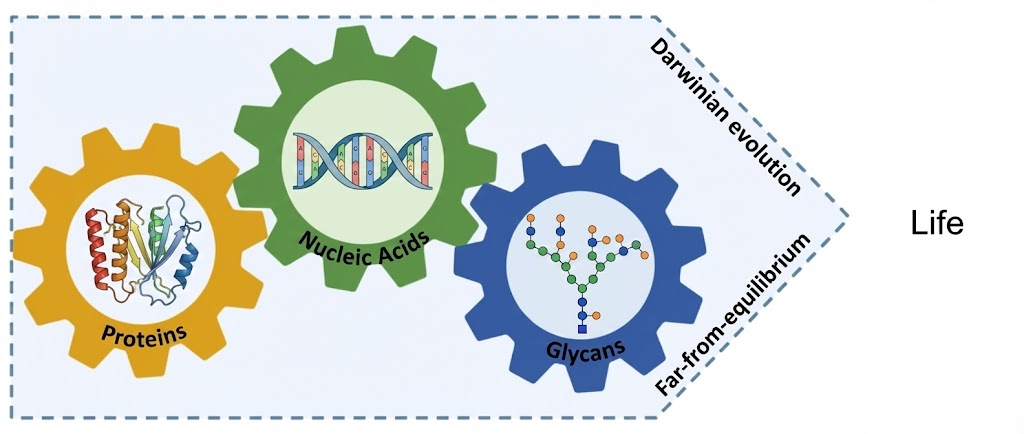
  • Glycans are made out of monosaccharides (e.g., glucose). 
  • Proteins are made out of amino acids.
  • Nucleic acids are made out of nucleotides

Most of our discussion will center on proteins and nucleic acids. Their constituents are themselves large, complex molecules. These monomers are built on an unchanging backbone and a variable side chain. For nucleotides, there are 4 available side chains (purine and pyrimidine nitrogenous bases). For amino acids, there are 20 available side chains (nonpolar, polar-uncharged, polar-positive, polar-negative R groups).

Organic polymers like polyesters aren’t particularly flexible. But biopolymers manifest extreme functional sophistication. And the polyfunctionality of nucleic acids and glycans must not be understated (Matange et al 2025).

Biopolymers also exhibit functional interdependence. Condensation of nucleotides is catalyzed by proteins, and condensation of amino acids is catalyzed by RNA. Biopolymers are heterocomplementary: proteins can recognize and bind to proteins, DNA, polyglycans, and small molecules. Nonbiological organic polymers do not manifest heterocomplementarity. 

LUCA from Microbiology

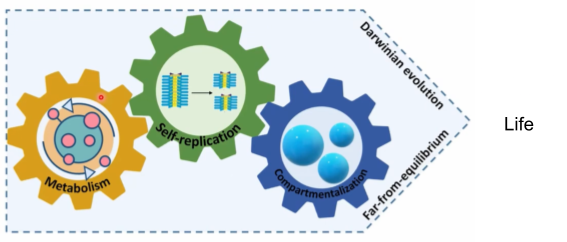
Another way to approach LUCA is to catalog biological universals: mechanisms shared by all life forms in existence. These universals can be organized in three categories: self-replication, metabolism, and compartmentalization. Let’s discuss each in turn.

The central genius of life was the discovery of proteins. By chaining amino acids together in long biopolymers, these polypeptides fold into arbitrary shapes. This invention is so powerful because in chemistry, structure determines function: arbitrary shapes unlock myriad potential behaviors. Proteins can form fibers, motors, containers, transporters, sensors, signals, optical devices, adhesives, pores, brushes, and pumps. 

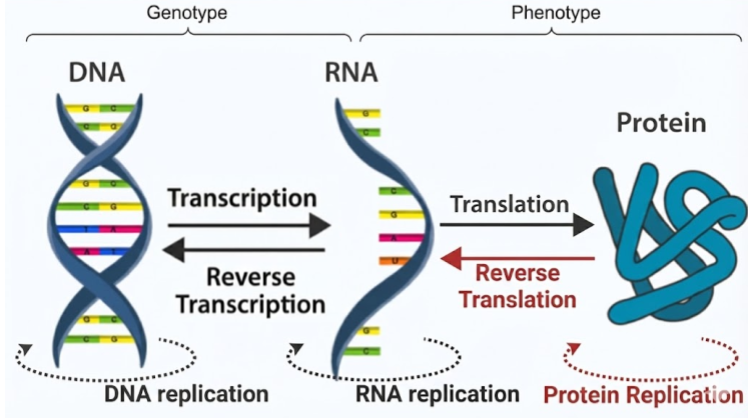
Proteins aren’t synthesized randomly – they are produced by template. Nucleotides combine together in long biopolymers known as nucleic acids. Double-stranded DNA is copied into single-stranded mRNA via the process of transcription. Then, during protein synthesis in the ribosome, codons (RNA triplets) specify which amino acids to attach in which order. This process of translation is defined by a codon-amino acid map known as the genetic code

A gene is simply a patch of RNA that completely specifies a particular protein sequence. Your genome (genotype) is nothing more than a recipe for a proteome (phenotype). When philosophers speak of an innate desire to survive, that motivation must be carried by proteins. Natural selection operates on protein design. Nothing more and nothing less.

The Central Dogma describes the flow of information in biopolymers. The black arrows are allowed processes; the red arrows are not observed. Proteins cannot replicate because they are incapable of base pairing. Once information gets into a protein, it cannot get out again!

Why is life so fixated on proteins? To better understand this, it helps to consider metabolism. This is a chemical reaction network which provides two basic functions: 

  1. Catabolism (“energy metabolism”), a destructive process which breaks down complex compounds. This prominently includes pathways of cellular respiration, a controlled version of combustion in which glucose is slowly converted into carbon dioxide, water, and energy. 
  2. Anabolism (“carbon metabolism”), a constructive process which biosynthesizes new organic compounds. This includes pathways of carbon fixation (like Acetyl-CoA), which converts simple inorganic carbon dioxide into complex organics.

Enzymes are protein-based catalysts that reduce the activation energy of thermodynamically favorable (exergonic) reactions, speeding them up by many orders of magnitude. Some metabolic steps, however, are thermodynamically disfavorable (endergonic) and will not proceed spontaneously. Enzymes handle these by coupling them to a favorable reaction, making the combined process exergonic. The enzyme then catalyzes that coupled reaction in the usual way — by lowering its activation energy.

The 20 amino acids have a limited chemical repertoire; cofactors extend it — enabling electron transfer, redox chemistry, and group transfers that amino acid side chains cannot perform alone. There are two categories of cofactors: metal ions (e.g., Mg²⁺, and Fe²⁺), and organic molecules (i.e., coenzymes), many of which are derived from B vitamins (e.g., NAD). Without its cofactor, an enzyme (called an apoenzyme) is typically inactive. The complete, functional assembly is the holoenzyme.

Finally, while the Central Dogma traditionally depicts a linear flow of information from DNA to RNA to Protein, metabolism reveals that this dogma is actually a loop. Proteins are not just passive end-products; they are the active machinery required to synthesize and maintain the very DNA and RNA that encode them, creating a self-sustaining cycle of synthesis and regulation.

Modern metabolism and self-replication does not occur in an undifferentiated “primordial soup”. They occur inside impermeable membranes made of phospholipids. 

Compartmentalization does a lot of heavy lifting at once: it shields nucleic acids from genetic parasites, buffers the system against environmental stress, and locally concentrates enzymes and substrates to speed reactions. By keeping metabolites and genetic material from simply diffusing away, it preserves hard-won products and information. Selective, protein-based gates regulate traffic—letting in foodstuffs, exporting waste—and can couple chemiosmosis to maintain a strong proton-motive force (≈3 pH units and ~200 mV), effectively a “cellular battery” that powers metabolic work. Finally, a stable compartment provides the physical unit that can reliably copy itself and propagate success via binary fission.

LUCA from Virology

Previous sections reconstructed LUCA as a cell. But modern oceans contain ten virions for every microbe. Viruses coevolve with cells, outnumber them, and may even predate them. Any reconstruction of early life that ignores them is incomplete.

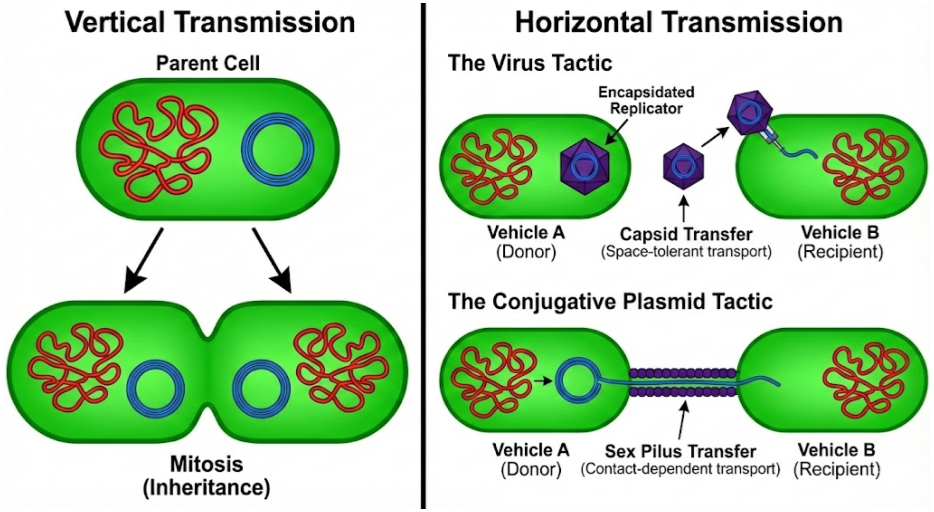
Once inside the cell, virulent viruses initiate the lytic cycle. Their genes hijack the host cell’s machinery to make more copies of themselves, ultimately rupturing the membrane and spilling newly-minted virions into the environment. They rely on horizontal transmission, in contrast to chromosomes which rely on vertical transmission via mitosis.  

Viruses are not the only mobile genetic element (MGE) using horizontal transmission. Cells can contain small (often circular) nucleic acid outside of the chromosomes. These plasmids don’t kill the cell to transfer their DNA, but instead use conjugative sex pili to copy themselves into neighboring cells. Plasmid genetic material can be more similar to viruses than the chromosome. Some plasmids and viruses differ by no more than a capsid gene (Krupovic & Bamford 2010).

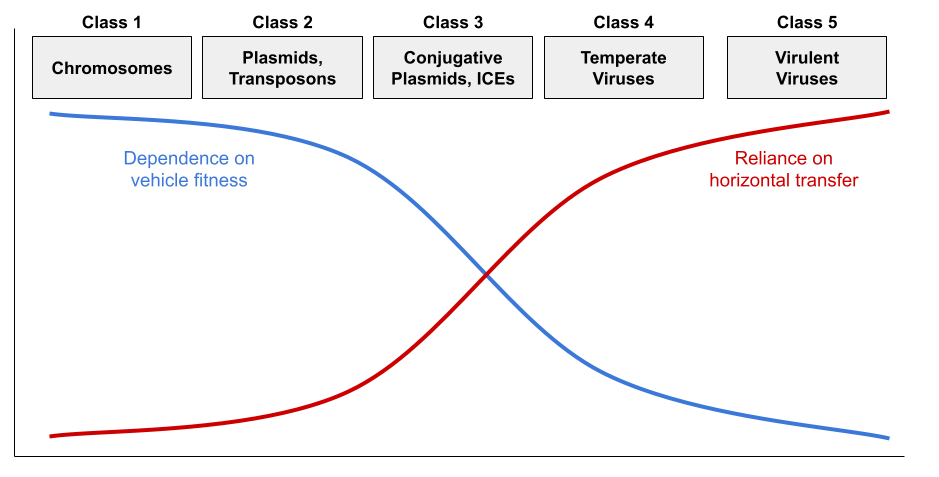
But virulence and conjugation are extremes. While virulent viruses are indifferent to cellular fitness, temperate viruses benefit from vertical transmission via the lysogenic cycle. They are thus less pathogenic. Some plasmids cannot construct the sex pilus and are capable of horizontal transmission only rarely. Jalasvuori (2012) shows that nucleic acids can be organized into a continuum:

This framework explains the distribution of replicator properties:

  • Class 3-5 all sense damage to the host cell, and initiate horizontal transfer when they detect vehicle stress. For example, herpesviruses reactivate from latency under host stress, producing cold sores.
  • Class 1 retains the core genome (e.g., DNA replication). Class 2-3 retain the accessory genome (e.g., virulence, antibiotic resistance).
  • Class 1-5 all retain addiction modules (a.k.a., MGE stabilization modules) like the toxin-antitoxin (TA) system (Mendoza-Guido & Rojas-Jimenez 2025). Many plasmids produce both a toxin and the antitoxin. Since the toxin has a longer half-life, loss of the plasmid entails death of the host. The restriction-modification (RM) system works in a very similar way (Kobayashi 2001).

There are three hypotheses for the origin of viruses:

  • The regression hypothesis claims that viruses are cells that progressively lost genetic information in their journey towards obligate parasitism.
  • The escape hypothesis claims that viruses originate from MGEs that gained increasingly sophisticated methods of horizontal transfer.
  • The virus-first hypothesis claims that viruses predate LUCA, originating from the primordial pool of replicators prior to the evolution of cells.

Dependent niches incentivize genome reduction. Mitochondria have transferred many genes to their host cells’ nuclei (Andersson & Kurland 1998), and parasitic bacteria like Rickettsia have shed much of theirs (Diop et al 2019). As obligate parasites, viruses should face the same reductive pressure — but what if they weren’t always parasitic? The discovery of giant viruses (in the phylum Nucleocytoviricota), with the first known translation-related machinery in a virus, was a revelation. In the spirit of the regression hypothesis, Boyer et al (2010) described them as living fossils of this fourth domain. But, while the relationship between reduction and parasitism is strong, recent phylogenetic work (Monttinen et al 2021) does not support this hypothesis.

The escape hypothesis accounts for some viral genes – particularly those encoding capsid proteins – which have clear cellular homologs. But viral hallmark genes, especially those involved in replication, are shared across diverse viral lineages yet absent from cellular genomes (Koonin et al 2006). If these genes escaped from cells, their cellular homologs should exist. Krupovic et al (2019) propose a chimeric origin that reconciles the escape and virus-first views. They argue that structural genes were captured from hosts, but replicator genes descend from primordial self-replicating elements that predate cells. 

Theoretical models reinforce this view — selfish genetic elements arise inevitably in any replicator system, suggesting that genetic parasites have been components of life from the very beginning. Before replicators evolved error correction mechanisms, early replicators also faced a hard physical constraint known as Eigen’s error threshold: RNA replication error rates impose an upper bound on genome size. Viroids, the simplest known replicators at ~300 nucleotides, sit near this boundary. If the chimeric model is correct and viral replicator genes descend from primordial self-replicating elements, viroids may be the closest living relatives of those elements. Their structural simplicity, their lack of protein-coding capacity, and the recent discovery of viroid-like circular RNAs across diverse environments (Lee et al 2023) are all consistent with deep ancestry.

Looking Beyond the Root

The portrait above shows LUCA as anaerobic, prokaryotic, protein-dependent, chromosomally organized, besieged by viruses. But every feature of that portrait is also a choice made from a menu of alternatives that were never taken:

  • Alternative codes. Of ~500 amino acid species, only 20 are included in the genetic code, even though codons could distinguish up to 64. Central metabolites like ornithine, GABA, and β-alanine are biosynthesized but never translated.
  • Alternative replicator backbones. 5-carbon sugars (ribose, deoxyribose) are a strange choice for a nucleic acid backbone (Eschenmoser 1999). Chemists have built alternative replicators from 4-carbon sugars (TNA) and 3-carbon sugars (GNA).
  • Alternative protein backbones. Proteins are built from α-amino acids, but foldamers made from β, γ, and δ-amino acids can be more resistant to proteolysis and heat denaturation than their α-counterparts (Gellman 1998).

McKay (2004) captures this puzzle visually: nonbiological chemistry produces smooth distributions of organic molecules, but life uses only a sparse set of spikes against that background.


Each of these alternatives works better than what life uses, on some axis. So the interesting question isn’t whether LUCA could have been different; it’s why it wasn’t. Why haven’t we found shadow biospheres running on TNA, or β-peptide proteomes, or 34-amino-acid codes? Two possibilities: either we haven’t looked hard enough, or these choices were frozen so early and so deeply that no later lineage could back out of them. Both answers require understanding what happened before LUCA: the evolution of the ribosome, the freezing of the code, the crystallization of a chromosome out of competing replicators.

Beyond the root of the tree of life lies the origin. That’s the subject of our next post.

References

  • Andersson & Kurland (1998). Reductive evolution of resident genomes
  • Bowman et al (2020). Root of the Tree: The Significance, Evolution, and Origins of the Ribosome
  • Boyer et al (2010). Phylogenetic and Phyletic Studies of Informational Genes in Genomes Highlight Existence of a 4th Domain of Life Including Giant Viruses
  • Caforio, A., et al. (2018). “Converting Escherichia coli into an archaebacterium with a hybrid heterochiral membrane.”
  • Diop et al (2019). Paradoxical evolution of rickettsial genomes
  • Eschenmoser (1999). Chemical etiology of nucleic acid structure.
  • Forterre (2006). The origin of viruses and their possible roles in major evolutionary transitions.
  • Forterre et al (2013) Origin and Evolution of DNA and DNA Replication Machineries
  • Gellman (1998). Foldamers: A manifesto.
  • Harris et al (2003). The genetic core of the universal ancestor. 
  • Jain et al (2014). Biosynthesis of archaeal membrane ether lipids
  • Jalasvuori (2012). Vehicles, replicators, and intercellular movement of genetic information: evolutionary dissection of a bacterial cell. 
  • Koonin et al (2006). The ancient virus world and evolution of cells. 
  • Kobayashi (2001). Behavior of restriction–modification systems as selfish mobile elements and their impact on genome evolution.
  • Krupovic et al (2019). Origin of viruses: primordial replicators recruiting capsids from hosts.
  • Krupovic & Bamford (2010). Order to the viral universe
  • Lee et al (2023). Mining metatranscriptomes reveals a vast world of viroid-like circular RNA
  • Matange et al (2025). Biological polymers: evolution, function, and significance.
  • McKay (2004). What Is Life—and How Do We Search for It in Other Worlds?
  • Mendoza-Guido & Rojas-Jimenez (2025). Beyond plasmid addiction: the role of toxin-antitoxin systems in the selfish behavior of mobile genetic elements. 
  • Monttinen et al (2021). The genomes of nucleocytoplasmic large DNA viruses: viral evolution writ large